Introduction
A complete comprehension of the Tagus River ecosystem is lacking. Due to the transboundary relevance of the Tagus River, its comprehension is paramount. Moreover, there is also insufficient information about the present and future impact posed by challenges in water management (Sondermann & de Oliveira, 2022), by climate change (Sondermann & de Oliveira, 2022), and anthropogenic alterations (Fernandes et al., 2020).
In terrestrial ecosystems most biomass comes from plants, where microbial biomass accounts for ~70% in the marine ecosystems (Bar-On et al., 2018). As rivers work as flows of nutrients and water it is relevant to get a clearer picture on the role of their microbiomes and phenotypes (Reddington et al., 2020; van Rossum et al., 2015). Since microorganisms have short lifecycles and large population size so their organization and structure correspond to a swift adaptation to the shifting environmental changes (Cavan et al., 2019; Collins et al., 2014; Hellweger et al., 2014). The global climate crisis is pressing the changes in marine ecosystems threatening the services they provide. This happens by major variations in the species present in marine habitats. To assess such fast shifts unified frameworks need to be developed that uses data from ecology, metabolism, and climate change (Baltar et al., 2019). Without this knowledge, the ability to understand, evaluate, and forecast the future of marine communities, ecosystems, and their services is almost inefficient.
In this Brief Report we wanted to elucidate the microbial profile of a coastline location in the Tagus River estuary (Beato, Lisboa), and infer the potential causes and consequences of the presence of the observed taxa.
Methods
Samples collection
The environmental water samples were collected in Praínha de Braço de Prata (Figure 1). Three water samples were collected using three sterile plastic containers (5 L) for microbiota analysis. The containers were submerged at 0.5 meters and opened below surface to guarantee no air-borne microorganisms’ collection.
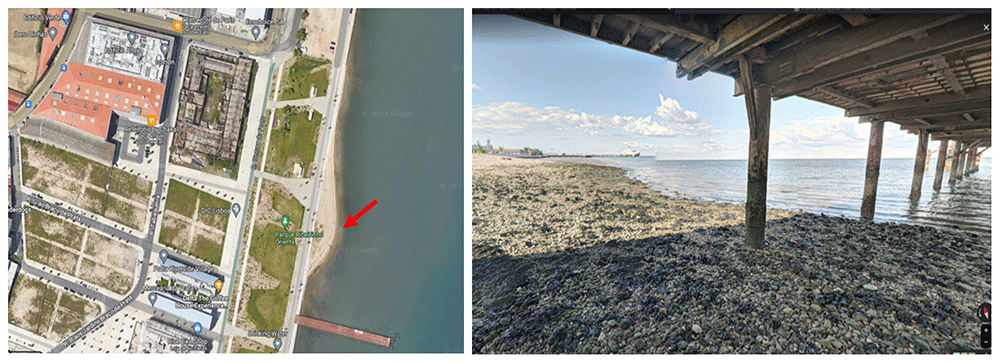
Figure 1. Sample collection site – ‘Praínha de Braço de Prata’.
Left: aerial view map of the site (red arrow). Right: photo of the site taken in May 2020 by Miguel Silva. Google Maps (n.d.) (View from Praínha de Braço de Prata). Retrieved in July 13, 2023, from https://www.google.com/maps/place/38°44'45.9%22N+9°05'47.2%22W/.
Total DNA was extracted from 5 L of each sample collected from the Tagus River and sequentially filtered through 20 μm, 3 μm, 0.45 μm, and 0.22 μm filters. The 0.45 μm and 0.22 μm filters were combined into a single 5 mL PowerWater Bead Pro Tube for DNA extraction using the DNeasy PowerWater Kit (Qiagen) according to the manufacturer's instructions. The purity of the extracted DNA was evaluated by measuring absorbance at 260 nm with a NanoDrop 1000 Spectrophotometer (Thermo Fisher Scientific, USA). The DNA was stored at −20 °C until further processing.
Amplification of then 16S rDNA gene and sequencing library preparation
For the rDNA gene amplification, Long Amp hot start Taq 2× master mix (New England Biolabs, MA, USA) was used at 1X along with 50 ng/μL of genomic DNA from Tagus River water samples. To amplify the full-length 16S rDNA bacterial gene, 0.25 μM of the primer pair 27F (5′-AGAGTTTGATCMTGGCTCAG-3′) and 1492R (5′-CGGTTACCTTGTTACGACTT-3′) were used. Both PCRs were conducted on a Biometra UNO II, using the following conditions for 16S rDNA gene: 1 cycle of 94 °C for 1 min, 35 cycles of 94 °C for 20 s, 55 °C for 30 s, and 65 °C for 2 min, and a final extension of 65 °C for 5 min. Subsequently, amplification products were visualized via gel electrophoresis and purified using the Solid Phase Reversible Immobilization (SPRI) technique with magnetic beads (Stotchevoi et al., 2020). The library was prepared from 200fmol input DNA from each sample using the Sequencing Native Barcoding Kit 24 V14 (SQK-NBD114.24) (Oxford Nanopore Technologies, Oxford, UK) in accordance with the manufacturer’s protocol.
Sequencing run
Sequencing runs were performed using R10.4.1 flow cells on a GridION sequencing platform and sequencing data were acquired in real time using MinKNOW 18.08.2 software. Sequencing data was stored in fastq files, with each file representing a batch of 4000 reads.
Bioinformatics analysis
Sequencing data was derived from 16S amplicons, with the exclusion of low-quality reads with the following parameters: a minimum quality score of 10, a minimum read length of 1200 bps, and the removal of reads with ambiguous bases (N's). The remaining reads underwent size selection, retaining those with lengths between 1200 bps and 1700 bps, accomplished through prinseq-lite (Schmieder & Edwards, 2011).
For taxonomic classification, we employed a Lowest Common Ancestor (LCA) approach. Clean reads were classified with Kraken version 2.1.2, using a reference database that includes a comprehensive collection of high-quality, annotated 16S rRNA sequences. The database used was the RefSeq plus GenBank Archaea and Bacteria database, created in May 2023. The classification process assigns each read to the lowest common ancestor of all genomes known to contain the specific sequence patterns identified in the read. This approach ensures robust taxonomic assignment by resolving ambiguities in read classification. For post-classification, the data underwent rarefaction and was subjected to several analyses: i) alpha-diversity group significance analysis (Bolyen et al., 2019); and ii) sample dissimilarity analysis - Principal Coordinates Analysis (PCoA): Applied for beta-diversity analysis based on the Bray-Curtis similarity index. Taxa abundance was determined, employing a genera prevalence cutoff of ≥ 0.01 (Bolyen et al., 2019). The sequencing depth for rarefaction was standardized to the sample with the lowest read count, which is 339,909 reads. This ensures that all samples are compared on an equal footing, avoiding biases introduced by differences in sequencing depth. In the rarefaction process, reads were randomly subsampled to match the sequencing depth of the sample with the lowest readcount (339,909 reads). This step was implemented to standardize the dataset while preserving the relative abundances of taxa within each sample. Then, data were subjected to determination of taxa abundance (with a genera prevalence cutoff of ≥ 0.01) (Bolyen et al., 2019) and Krona Plots (by hierarchical order of Operational Taxonomic Units frequency).
The validation of our bioinformatics approach was performed using the ZymoBIOMICS™ Microbial Community Standard. Specifically, we sequenced this mock community using 16S rRNA amplicon sequencing and analyzed the data with the same pipeline applied to our samples. The proportions of sequencing reads classified to each taxon closely matched the expected proportions in the mock community. This agreement between the observed and expected proportions confirms the accuracy of our methodology in reflecting community composition, thereby validating the reliability of our approach for the samples studied.
For post-classification, the data underwent rarefaction and was subjected to several analyses: i) alpha-diversity group significance analysis (Bolyen et al., 2019); and ii) sample dissimilarity analysis - Principal Coordinates Analysis (PCoA): Applied for beta-diversity analysis based on the Bray-Curtis similarity index. Taxa abundance was determined, employing a genera prevalence cutoff of ≥ 0.01 (Bolyen et al., 2019). The sequencing depth for rarefaction was standardized to the sample with the lowest read count, which is 339,909 reads. This ensures that all samples are compared on an equal footing, avoiding biases introduced by differences in sequencing depth. In the rarefaction process reads were randomly subsampled to match the sequencing depth of the sample with the lowest read count (339,909 reads). This step was implemented to standardize the dataset while preserving the relative abundances of taxa within each sample.
Results
Nanopore sequencing general output
The three samples showed similar quality parameters which gives confidence that the sample collection and the sequencing approach were appropriately performed (Table 1). Specifically, the mean read length is equal in the three analyses (1,570-1,574 bases), the same for the mean read quality (Q = 17.7-18.0), and the percentage of reads above the cutoffs. The sample 2 produced less reads (and bases) than the other two samples, but that did not hamper the quality of the output.
Table 1. Main nanopore sequencing quality parameters from the three environmental samples.
| Quality feature (units) | Sample 1 | Sample 2 | Sample 3 |
|---|
| Total number of reads (thousand) | >534 | >339 | >474 |
| Total number of classified reads (%) | 99.82 | 99.87 | 99.91 |
| Total number of bases (million) | >841 | >533 | >746 |
| Mean read length (bases) | 1,574 | 1,570 | 1,571 |
| Read length N50 (bases) | 1,594 | 1,584 | 1,582 |
| Mean read quality (Q score)* | 17.5 | 18.0 | 18.0 |
| Reads above quality 7 cutoff (%) | 100.0 | 100.0 | 100.0 |
| Reads above quality 10 cutoff (%) | 100.0 | 100.0 | 100.0 |
| Reads above quality 15 cutoff (%) | 84.1 | 84.3 | 84.7 |
Prainha de Braço de Prata microbiota
The top main phylum, genus, and species observed in the three water samples are shown in Table 2.
Table 2. Three top phylum, genera, and species identified in the three samples with their respective relative abundance.
| Sample | Rank | Taxa (relative abundance, %) |
|---|
| 1 | Phylum | Pseudomonadota (36), Actinomycetota (10), Bacteroidota (11) |
| Family | Flavobacteriaceae (10), Roseobacteraceae (10), Egibacteraceae (7) |
| Genus | Egibacter (7), Tateyamaria (6), Ralstonia (6) |
| Species | Egibacter rhizosphaerae (7), Tateyamaria omphalii (6), Ralstonia solanacearum (6) |
| 2 | Phylum | Pseudomonadota (65), Actinomycetota (15), Bacteroidota (15) |
| Family | Burkholderiaceae (14), Roseobacteraceae (14), Flavobacteriaceae (13) |
| Genus | Ralstonia (14), Candidatus Pelagibacter (11), Egibacter (10) |
| Species | Ralstonia solanacearum (14), Candidatus Pelagibacter sp. (11), Egibacter rhizosphaerae (10) |
| 3 | Phylum | Pseudomonadota (45), Actinomycetota (13), Bacteroidota (10) |
| Family | Pelagibacteraceae (17), Egibacteraceae (10), Flavobacteriaceae (9) |
| Genus | Candidatus Pelagibacter (17), Egibacter (10), Ralstonia (7) |
| Species | Uncultivated bacterium from the SAR86 clade (16), Ralstonia solanacearum (10), Egibacter
rhizosphaerae (7) |
In order to assess the Praínha do Braço de Prata microbiota we qualified and quantified the shared taxa between the three samples (Figure 2).
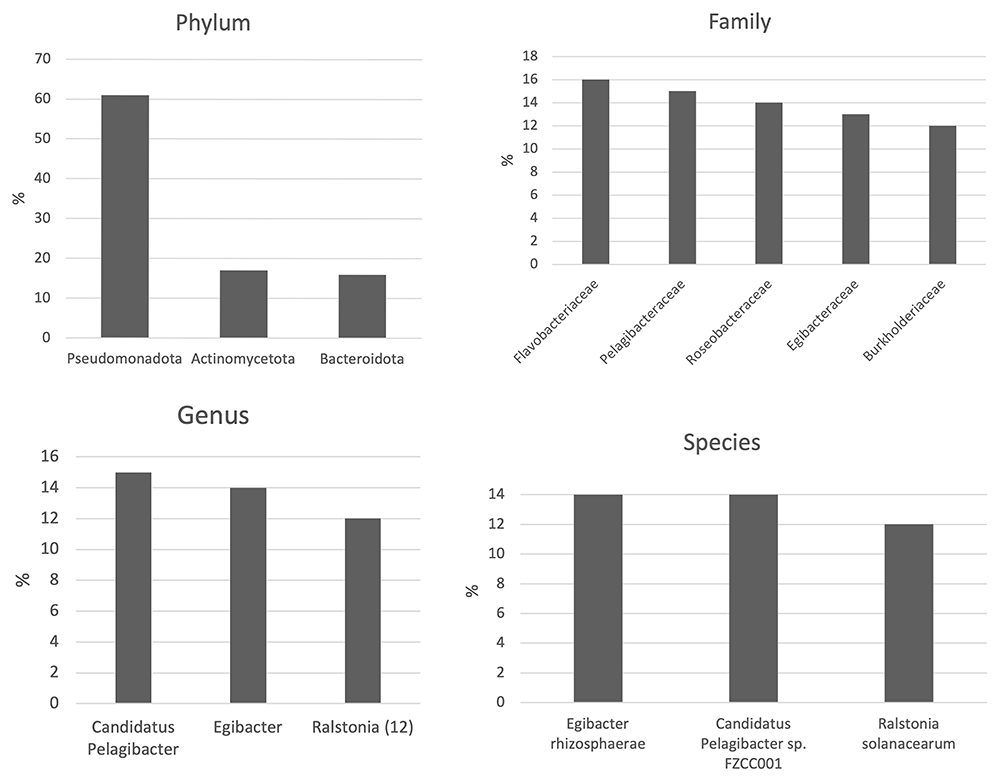
Figure 2. Shared taxa at the phylum, family, genus, and species level between the three samples.
Only the taxa with relative abundances above 10% are sgown.
The microbial profiles differed between sample 1 and sample 3 at the phylum, genus, and species level (Shannon index; p-value < 0.05). The sample 1 and sample 2 showed similar alpha-diversities (Shannon index; p-value > 0.05). The samples showed incongruent dissimilarities microbial profiles amongst them. There were no differences amongst the three samples’ microbial profiles at the genus level. At the species level, the microbial profile from sample 3 is different from sample 1 and 2, but sample’s 2 and sample’s 1 microbial profile are similar. Lastly, he phylum level, the microbial profile from sample 2 is different from sample 1 and 3, and sample’s 1 shows a similar microbial profile than sample 3 (PERMANOVA R2 = 1).
Ecosystem services
Based on the shared taxa found between the three samples we used the top three genus and species to assess in PubMed database studies that inform the potential ecosystem services these taxa might be performing in the Tagus River. In total it was identified 17 articles related to Pelagibacter gen. and Pelagibacter sp., and 37 related to Ralstonia gen. and Ralstonia solanacearum. No article was found for Egibacter or Egibacter rhizosphaerae. Additionally, no review article was published about the ecosystem services of these taxa in the defined period of search in the systematic analysis. The relevant articles found follow as: 14 for Pelagibacter gen. and Pelagibacter sp., and 11 for Ralstonia gen. and R. solanacearum (Table 3).
Table 3. Relevant ecosystem services present in literature for the most abundant taxa.
| TaxaID | Relevant reference | Ecosystem service/ phenotype |
|---|
Candidatus
Pelagibacter or
SAR11 | Rivers et al., 2013 | Re-establishing the pre-oil spill microbial community structure. |
| Tsementzi et al., 2019 | Oligotrophy; auxotrophy for reduced sulfur (lacking a sulfite oxidase); differential phage
predation and/or adaptation (fine tuning) of core genes to environmental conditions (e.g.
temperature). |
| Liu et al., 2023 | Resistance to salinity; ability to grow in low nutrients sites (like dissolved inorganic nitrogen);
utilizers of dissolved organic carbon. |
| Han et al., 2020 | Heterotrophy; carbon cycling; generalista; dimethylsulfoniopropionate degradation. |
| Cerro-Gálvez et al., 2021 | Oligotrophy; antitoxicity strategies (to anthropogenic dissolved organic carbon; high tolerance
to hydrophobic chemicals |
| Chun et al., 2021 | ‘Skeleton’-formation and participate in species-engineered microenvironments; dominant in
surface waters under stratified summer conditions; fine-tuning microbial modular structures;
present in the nanoparticle-associated fraction; nitrate reduction, nitrification, dark sulfide
oxidation. |
| Delpech et al., 2021 | Oligotrophy; follow; abundant in low nutrient concentrations and in stratified surface waters
phytoplankton blooms |
| Sun et al., 2021 | Able to maximize nutrients uptake even when in low concentrations. |
| Zhang et al., 2022 | Present in anticyclonic eddies, in warmer surface water temperatures, where there is less
diverse bacterial communities; oligotrophy; survives at lower salinity and chlorophyl a
concentrations but tolerate higher concentrations. |
| Anderson & Harvey, 2022 | Oligotrophy; rely on copiotrophs to assimilate sources of carbon and other metabolites. |
| Fortin et al., 2022 | Predominant in Alexandrium sp. blooms; present in the particle-attached fraction of the surface
water. |
| Guo et al., 2022 | Involved in nitrogen and/ or sulfur cycling: prevalent in oxygen-rich surface oceans |
| Larkin et al., 2023 | Heterotrophy; partition marine niche spaces; adapted to regions with low inorganic nitrogen
supply; reveals habitat-specific partitioning in replication strategies for surface ocean
communities |
| Sun et al., 2023 | Prefers nitrogen-containing carbohydrates, oligo-glucans, oligo-cellulose, and lignin-derived
aromatic fragments; polysaccharide degradation; scavenging mode in free-living fractions for
the assimilation of dissolved carbohydrates, oligotrophy; key in significant flux of dissolved
organic carbon and nutrient mineralization at the bottom of watery ecosystems. |
| Ralstonia | Coll et al., 2020 | Degradation of contaminants (e.g., organic pollutants); |
| Rasool et al., 2020 | Tolerance to heavy metals like thallium; role in distributing metals; dominant in midstream soil
samples; tent to enrich in bank of steam soils |
| Chang et al., 2021 | Rare pathogenicity and transmitted through polluted waters. |
| Liu et al., 2023 | Degrade microcystins; may enhance host-resistance to toxic cyanobacterial stress by affecting
the whole bacterial community. |
| Yang et al., 2023a | May utilize diesel-derived organic intermediates produced by hydrocarbon degraders. |
| Yang et al., 2023b | Thrive in soil discharge and plant decomposition conditions; decomposition of several aromatic
hydrocarbons. |
| R. solanacearum | Planas-Marquès
et al., 2020 | Vascular pathogen; wide geographic distribution; colonizes many hosts, persists in soil and
water for long periods; moves horizontally through the xylem ring by degrading the cell wall of
primary xylem or pit membranes in the secondary xylem vessels. |
| Biosca et al., 2021 | Infects ornamental plants; lytic lifestyle. |
| Wen et al., 2023 | Cause wilt disease; affects the composition and diversity of rhizosphere microbiome |
Conclusion
Rivers present a key importance for human life and for nature balance since it is the primordial source of renewable water (Vörösmarty et al., 2010). However, there is a lack of studies that focus the microbial diversity in rivers compared to marine or lake ecosystems that could elucidate about its relevance for urban areas and ecosystems services. The dynamic flow of water and nutrients in rivers make their study complex but necessary to answer foundational questions regarding land use and novel threats like antimicrobial resistance and the emergence of invasive species (Reddington et al., 2020; van Rossum et al., 2015). Despite the low number of samples analysed through this Brief Report, it is clear that we have insufficient information about the taxonomy of the most abundant taxa detected like Pelagibacter and Egibacter and more research is necessary to shed light on their relevant phenotypes aiming a better comprehension of the Tagus River ecosystem. Interestingly, the most abundant Pelagibacter strains were commonly found in marine waters rather than freshwaters (“FZCC0015” and “IMCC9063”). This observation constitutes a finding that deserves better attention since it may represent evidence of salinity changes in the Tagus River estuary.
Ethics and consent
Ethical approval and consent were not required.
Contributorship CRediT statement
Cristiano Pedroso-Roussado: Conceptualization, Methodology, Investigation, Visualization, Writing – Original Draft, Writing – Review & Editing. Mariana Pestana: Validation, Writing – Review & Editing, Funding acquisition. Ricardo Dias: Methodology, Investigation, Data curation, Visualization. Mónica Nunes: Methodology, Investigation, Data curation, Visualization. Pedro Pascoal: Methodology, Investigation, Data curation, Visualization. Marcelo Pereira: Methodology, Investigation, Data curation, Visualization. Nuno Nunes: Validation, Writing – Review & Editing, Supervision.
Faculty Opinions recommendedReferences
- Anderson SR, Harvey EL:
Estuarine microbial networks and relationships vary between environmentally distinct communities.
PeerJ.
2022; 10: e14005. PubMed Abstract
| Free Full Text
- Baltar F, Bayer B, Bednarsek N, et al.:
Towards integrating evolution, metabolism, and climate change studies of marine ecosystems.
Trends Ecol Evol.
2019; 34(11): 1022–1033. PubMed Abstract
| Publisher Full Text
- Bar-On YM, Phillips R, Milo R:
The biomass distribution on Earth.
Proc Natl Acad Sci U S A.
2018; 115(25): 6506–6511. PubMed Abstract
| Publisher Full Text
| Free Full Text
- Biosca EG, Català-Senent JF, Figàs-Segura À, et al.:
Genomic analysis of the first European bacteriophages with depolymerase activity and biocontrol efficacy against the phytopathogen Ralstonia solanacearum.
Viruses.
2021; 13(12): 2539. PubMed Abstract
| Publisher Full Text
| Free Full Text
- Bolyen E, Rideout JR, Dillon MR, et al.:
Reproducible, interactive, scalable and extensible microbiome data science using QIIME 2.
Nat Biotechnol.
2019; 37(8): 852–857. PubMed Abstract
| Publisher Full Text
| Free Full Text
- Cavan EL, Henson SA, Boyd PW:
The sensitivity of subsurface microbes to ocean warming accentuates future declines in particulate carbon export.
Front Ecol Evol.
2019; 6: 230. Publisher Full Text
- Cerro-Gálvez E, Dachs J, Lundin D, et al.:
Responses of coastal marine microbiomes exposed to Anthropogenic Dissolved Organic Carbon.
Environ Sci Technol.
2021; 55(14): 9609–9621. PubMed Abstract
| Publisher Full Text
| Free Full Text
- Chang CJ, Zhang J, Tsai YL, et al.:
Compositional features of distinct microbiota base on serum extracellular vesicle metagenomics analysis in moderate to severe psoriasis patients.
Cells.
2021; 10(9): 2349. PubMed Abstract
| Publisher Full Text
| Free Full Text
- Chun SJ, Cui Y, Baek SH, et al.:
Seasonal succession of microbes in different size-fractions and their modular structures determined by both macro- and micro-environmental filtering in dynamic coastal waters.
Sci Total Environ.
2021; 784: 147046. PubMed Abstract
| Publisher Full Text
- Coll C, Bier R, Li Z, et al.:
Association between aquatic micropollutant dissipation and river sediment bacterial communities.
Environ Sci Technol.
2020; 54(22): 14380–14392. PubMed Abstract
| Publisher Full Text
| Free Full Text
- Collins S, Rost B, Rynearson TA:
Evolutionary potential of marine phytoplankton under ocean acidification.
Evol Appl.
2014; 7(1): 140–155. PubMed Abstract
| Publisher Full Text
| Free Full Text
- Delpech LM, Vonnahme TR, McGovern M, et al.:
Terrestrial inputs shape coastal bacterial and archaeal communities in a high Arctic fjord (Isfjorden, Svalbard).
Front Microbiol.
2021; 12: 614634. PubMed Abstract
| Publisher Full Text
| Free Full Text
- Fernandes MR, Aguiar FC, Martins MJ, et al.:
Long-term human-generated alterations of Tagus river: effects of hydrological regulation and land-use changes in distinct river zones.
CATENA.
2020; 188: 104466. Publisher Full Text
- Fortin SG, Song B, Anderson IC, et al.:
Blooms of the harmful algae Margalefidinium polykrikoides and Alexandrium monilatum alter the York River Estuary microbiome.
Harmful Algae.
2022; 114: 102216. PubMed Abstract
| Publisher Full Text
- Guo R, Ma X, Zhang J, et al.:
Microbial community structures and important taxa across oxygen gradients in the Andaman Sea and eastern Bay of Bengal epipelagic waters.
Front Microbiol.
2022; 13: 1041521. PubMed Abstract
| Publisher Full Text
| Free Full Text
- Han D, Kang HY, Kang CK, et al.:
Seasonal mixing-driven system in estuarine–coastal zone triggers an ecological shift in bacterial assemblages involved in phytoplankton-derived DMSP degradation.
Microb Ecol.
2020; 79(1): 12–20. PubMed Abstract
| Publisher Full Text
- Hellweger FL, van Sebille E, Fredrick ND:
Biogeographic patterns in ocean microbes emerge in a neutral agent-based model.
Science.
2014; 345(6202): 1346–1349. PubMed Abstract
| Publisher Full Text
- Larkin AA, Hagstrom GI, Brock ML, et al.:
Basin-scale biogeography of prochlorococcus and SAR11 ecotype replication.
ISME J.
2023; 17(2): 185–194. PubMed Abstract
| Publisher Full Text
| Free Full Text
- Liu H, Yang X, Yang W, et al.:
Gut microbiota of freshwater gastropod (Bellamya aeruginosa) assist the adaptation of host to toxic cyanobacterial stress.
Toxins (Basel).
2023; 15(4): 252. PubMed Abstract
| Publisher Full Text
| Free Full Text
- Pedroso-Roussado C, Pestana M, Dias R, et al.:
Tagus river bacterial microbiota found in praina do braco de prata.
Sequence Read Archiveı.
2024; Retrieved July 4, 2024. https://www.ncbi.nlm.nih.gov/bioproject/PRJNA1116853/
- Planas-Marquès M, Kressin JP, Kashyap A, et al.:
Four bottlenecks restrict colonization and invasion by the pathogen Ralstonia solanacearum in resistant tomato.
J Exp Bot.
2020; 71(6): 2157–2171. PubMed Abstract
| Publisher Full Text
| Free Full Text
- Rasool A, Nasim W, Xiao T, et al.:
Microbial diversity response in thallium polluted riverbank soils of the Lanmuchang.
Ecotoxicol Environ Saf.
2020; 187: 109854. PubMed Abstract
| Publisher Full Text
- Reddington K, Eccles D, O’Grady J, et al.:
Metagenomic analysis of planktonic riverine microbial consortia using nanopore sequencing reveals insight into river microbe taxonomy and function.
GigaScience.
2020; 9(6): giaa053. PubMed Abstract
| Publisher Full Text
| Free Full Text
- Rivers AR, Sharma S, Tringe SG, et al.:
Transcriptional response of bathypelagic marine bacterioplankton to the Deepwater Horizon oil spill.
ISME J.
2013; 7(12): 2315–2329. PubMed Abstract
| Publisher Full Text
| Free Full Text
- Schmieder R, Edwards R:
Quality control and preprocessing of metagenomic datasets.
Bioinformatics.
2011; 27(6): 863–864. PubMed Abstract
| Publisher Full Text
| Free Full Text
- Sondermann MN, de Oliveira RP:
Climate adaptation needs to reduce water scarcity vulnerability in the Tagus River basin.
Water.
2022; 14(16): 2527. Publisher Full Text
- Sun FL, Wang YS, Wu ML, et al.:
Bacterial community variations in the South China Sea driven by different chemical conditions.
Ecotoxicology.
2021; 30(9): 1808–1815. PubMed Abstract
| Publisher Full Text
- Sun CC, Zhao WJ, Yue WZ, et al.:
Polymeric carbohydrates utilization separates microbiomes into niches: insights into the diversity of microbial carbohydrate-active enzymes in the inner shelf of the Pearl River Estuary, China.
Front Microbiol.
2023; 14: 1180321. PubMed Abstract
| Publisher Full Text
| Free Full Text
- Tsementzi D, Rodriguez-R LM, Ruiz-Perez CA, et al.:
Ecogenomic characterization of widespread, closely-related SAR11 clades of the freshwater genus “Candidatus Fonsibacter” and proposal of Ca. Fonsibacter lacus sp. nov.
Syst Appl Microbiol.
2019; 42(4): 495–505. PubMed Abstract
| Publisher Full Text
- Van Rossum T, Peabody MA, Uyaguari-Diaz MI, et al.:
Year-long metagenomic study of river microbiomes across land use and water quality.
Front Microbiol.
2015; 6: 1405. PubMed Abstract
| Publisher Full Text
| Free Full Text
- Vörösmarty CJ, McIntyre PB, Gessner MO, et al.:
Global threats to human water security and river biodiversity.
Nature.
2010; 467(7315): 555–561. PubMed Abstract
| Publisher Full Text
- Wen T, Xie P, Liu H, et al.:
Tapping the rhizosphere metabolites for the prebiotic control of soil-borne bacterial wilt disease.
Nat Commun.
2023; 14(1): 4497. PubMed Abstract
| Publisher Full Text
| Free Full Text
- Yang R, Peng C, Ye Y, et al.:
Succession patterns of microbial composition and activity following the diesel spill in an urban river.
Microorganisms.
2023b; 11(3): 698. PubMed Abstract
| Publisher Full Text
| Free Full Text
- Yang M, Wen G, Cao S, et al.:
The formation of Double Metalimnetic Oxygen Minima in a drinking water reservoir and its influence on bacterial community.
Sci Total Environ.
2023a; 860: 160540. PubMed Abstract
| Publisher Full Text
- Zhang W, Yuan W, Chen L, et al.:
Uniqueness and dependence of bacterial communities on microplastics: comparison with water, sediment, and soil.
Microb Ecol.
2022; 84(4): 985–995. PubMed Abstract
| Publisher Full Text