1 Introduction
Modern industrial, agricultural, and urban activities release a continually expanding volume and variety of contaminants into the environment. Pervasive infiltration, increasingly complex pollutant mixtures, and a growing catalogue of emerging 'chemicals of concern' now burden soil ecosystems (Brühl et al., 2024) demanding robust Ecological Risk Assessment (ERA) frameworks that address these complexities with statistical rigor and harness new technologies to identify and interpret sensitive risk indicators. Early ERA relied on toxicity benchmarks established by exposing standardized organisms to contaminants in artificial media and assessing acute response. Because these chemical concentration thresholds ignore site-specific bioavailability and the in situ interactions that govern actual contaminant effects, they misrepresent the actual risks present under field conditions (Chapman, 1990; Posthuma et al., 2002; Environment Agency, UK, 2022).
The TRIAD ERA framework, originally developed as the Sediment Quality Triad, by Long and Chapman (1985), overcomes this limitation by integrating chemical, ecotoxicological, and ecological data as three distinct Lines of Evidence (LoEs) in a site-specific assessment. Within each LoE, the measured endpoints are converted into scores, weighted by criteria reflecting their reliability and ecological relevance, and integrated; the LoEs are then also weighted and combined into a single integrated risk (IR) value (Chapman, 1986; Chapman, 1997; Jensen and Mesman, 2006; Dagnino et al., 2008). The framework follows a three-tiered structure, with each tier adding more detailed tests and sophisticated analysis methods, increasing time and cost, to resolve assessment uncertainty remaining from the previous tier.
Figure 1 shows tests and measured parameters commonly applied at each tier of the TRIAD framework (reproduced for reference from Part I).
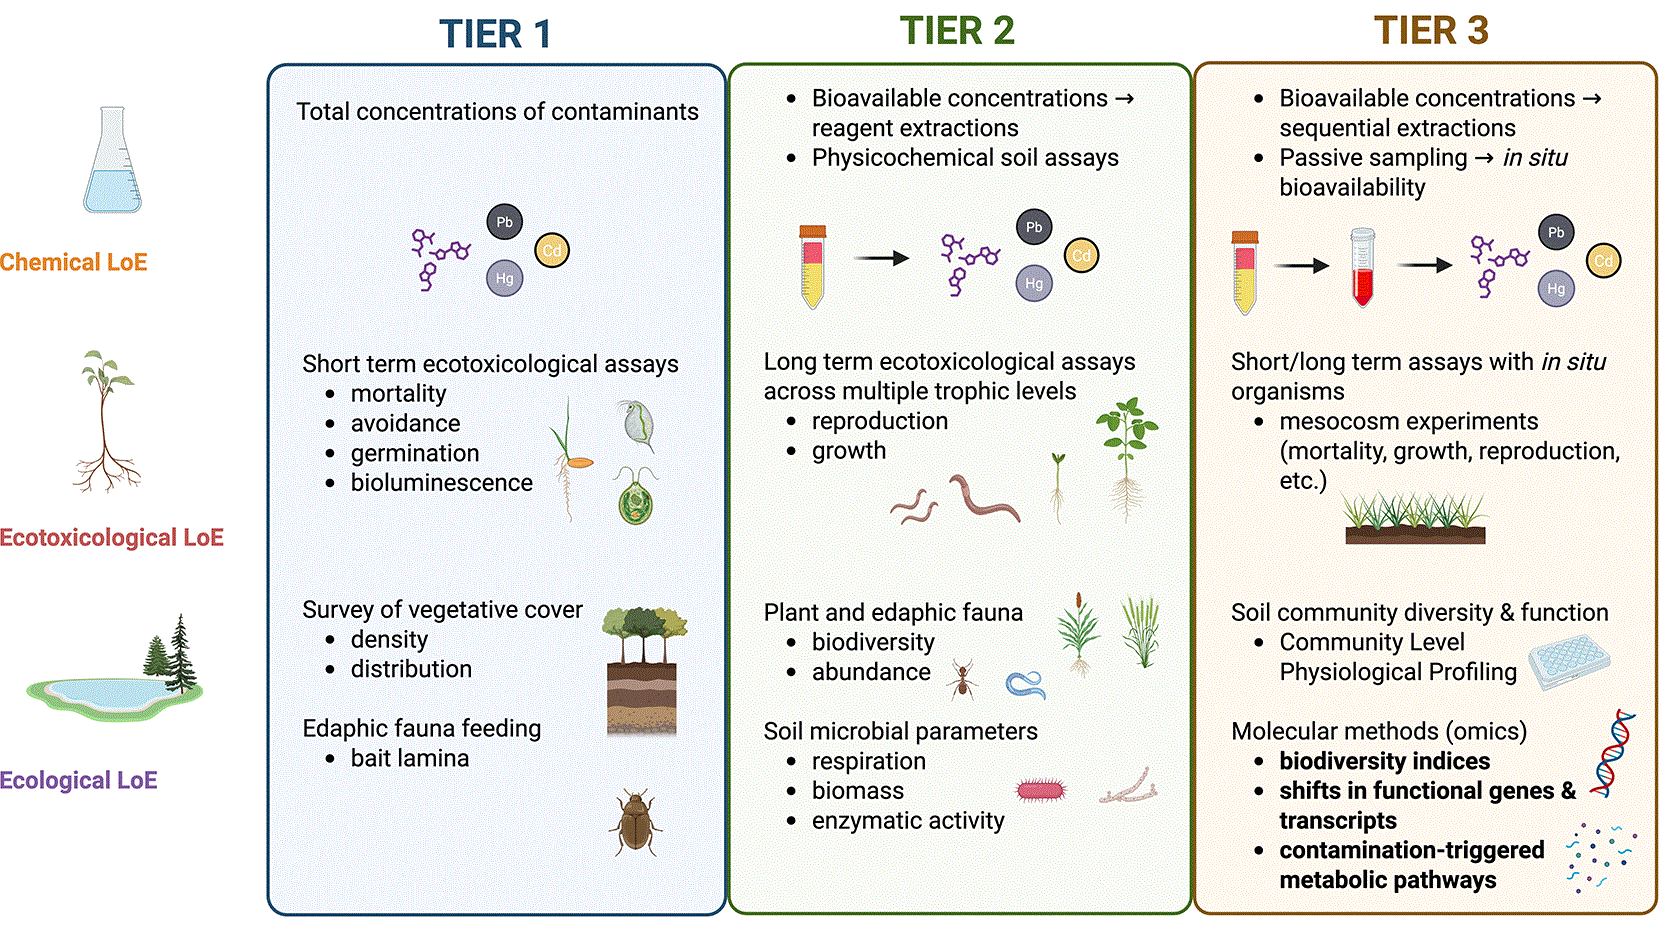
Figure 1. Common tests and measured parameters currently used in the TRIAD, organized by tier and by chemical, ecotoxicological, and ecological Lines of Evidence.
Each successive tier resolves residual uncertainty from the previous one by adding more targeted and sensitive methods, capturing both acute and chronic effects. Molecular omic methods (bold) currently enter at tier 3 as high-resolution indicators of community diversity and function; however, they have rarely been applied in TRIAD-based assessments (Grassi et al., 2022). This work examines a restructuring of the current scheme to expand the role of omic methods. Adapted from Part I (Djomehri et al., 2026). Created with BioRender.
Part I (Djomehri et al., 2026) establishes the analytical and decision-support components of a modernized TRIAD framework, including structured expert judgement, MCDA, multivariate analyses, machine learning and minimum datasets of soil quality indicators. These developments improve transparency, uncertainty tracking and analytical rigor in TRIAD assessments, while providing a more robust basis for indicator selection, weighting and data integration.
While the analytical advances proposed in Part I (Djomehri et al., 2026) improve the transparency and applicability of the TRIAD framework, they cannot extract high-resolution insight from coarse data, remaining fundamentally bound by the sensitivity of the available response parameters. The growing understanding that soil functionality underpins multiple ecosystem services has driven widespread criticism of the lack of sensitive biological parameters in terrestrial ERA (Barrios, 2007; Faber and Van Wensem, 2012; Stone et al., 2016; Bünemann et al., 2018). Within TRIAD, this shortcoming weakens the ecological LoE in particular, the line that carries the most site-specific information and should therefore weigh most heavily in assessment (Chapman and Hollert, 2006; Dagnino et al., 2008). Including sub-lethal microbial indicators, such as enzymatic activity, cellular respiration, microbial biomass, and community-level physiological profiling, in higher-tiered assessments has partially addressed this gap, providing data on contaminant bioavailability, toxicity, and functional diversity unattainable through conventional ecological surveys (Bastida et al., 2008; Aponte et al., 2020; Klauberg-Filho et al., 2023).
The advancement of molecular methods now offers a level of detection which surpasses these traditional microbial parameters. ‘Omic' techniques (genomics, transcriptomics, proteomics, and metabolomics) probe genes, proteins, and metabolites directly from environmental samples, capturing the effects of contamination at the molecular level (Bouchez et al., 2016). Indicator-selection studies for ERA point to molecular microbial measures (Ritz et al., 2009). In a structured assessment of biological indicators for European soil monitoring, seven of the ten top-ranked indicators used molecular methods, with community diversity and functional gene measures scoring highest for discrimination, multifunctionality, and responsivity (Stone et al. 2016). A growing body of literature advocates incorporating omic technologies to address the scarcity of sensitive ecological endpoints (Bouchez et al., 2016; Bünemann et al., 2018; Djemiel et al., 2022).
Omic methods profile biological response to contamination at two scales. At the community scale, meta-omic approaches characterize the taxa, functional genes, expressed pathways, proteins, and metabolites of whole, uncultured environmental samples (Segata et al., 2013), supplying high-resolution indicators for the ecological LoE. At the organism and population scale, they reveal the molecular stress responses and toxicity mechanisms within individuals and species such as plants, earthworms, and other soil fauna, providing sensitive indicators for both the ecotoxicological and ecological LoEs. Across both scales, different methods read distinct molecular layers. Targeted high-throughput DNA sequencing gauges the relative abundance of taxa and functional genes, exposing contamination-induced shifts in community composition and specified functional potential. High-density microarrays read the same targeted layer using probes built from known reference sequences, which bind and quantify their target genes by hybridization, rather than sequencing the targets themselves. Metagenomic sequencing is untargeted, capturing broader patterns in genetic response. RNA sequencing measures the genes actually expressed, detecting the activation of stress-response pathways and the disruption of essential functions under contamination. Proteomic and metabolomic analyses examine the biochemical layer, registering the up- or down-regulation of proteins and metabolites triggered by contaminant exposure. Integrating these layers through multi-omics traces the path from gene presence and expression to the proteins and metabolites produced, providing the most detailed picture of the mechanisms by which contamination drives toxicity.
Figure 2 summarizes the molecular risk bioindicators these methods yield for soil contamination assessment.
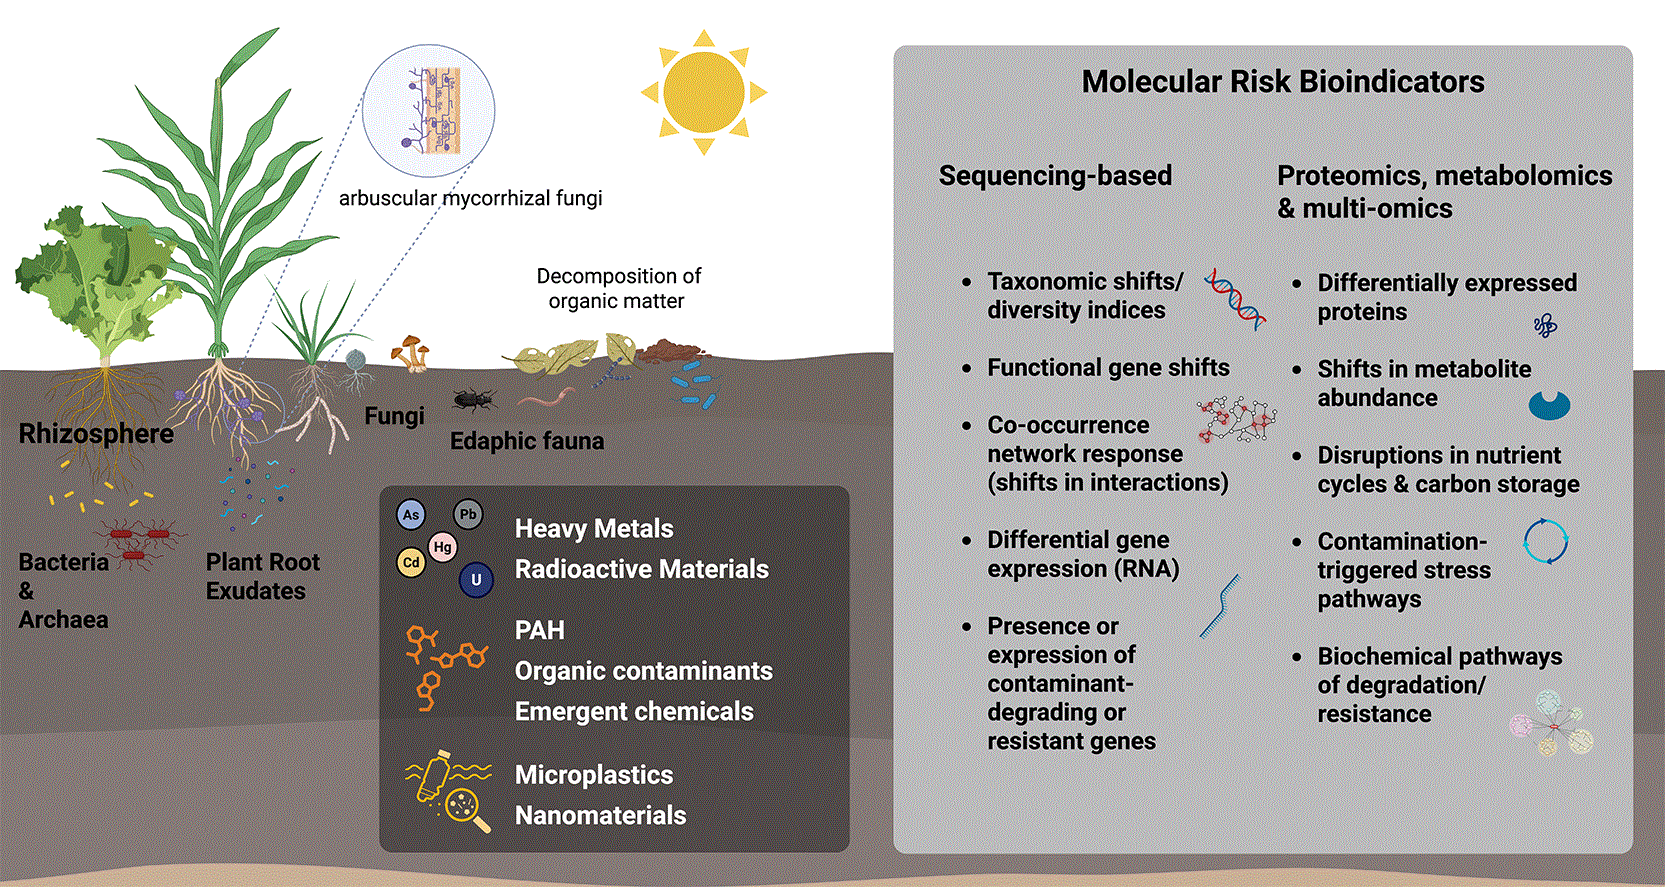
Figure 2. A representation of molecular risk bioindicators of soil contamination derived from omic methods.
Soil organisms - plant roots and their rhizosphere, bacteria, archaea, fungi, and edaphic fauna - are exposed to diverse contaminants, including heavy metals, radioactive materials, organic pollutants, and emerging chemicals such as microplastics and nanomaterials. DNA/RNA sequencing and proteomic, metabolomic, and multi-omic analyses translate the biological response to this exposure into measurable indicators of ecological risk, spanning taxonomic and functional gene shifts, altered gene expression, and changes in proteins, metabolites, and the biochemical pathways underlying contaminant degradation, resistance, and toxicity. Created with BioRender.
Beyond their sensitivity, molecular indicators offer several advantages that justify their inclusion in the modernized TRIAD framework. A major strength is their ability to identify the source of an observed effect: separating contamination from co-occurring abiotic stressors, distinguishing individual contaminant signatures within complex mixtures, and detecting synergistic or antagonistic interactions between compounds or climatic factors (You et al., 2022; Silva et al., 2024). Integrated multi-omics can expose both a contaminant's immediate toxic effects and indirect yet significant harm transmitted through altered soil properties or disrupted microbial communities (Han et al., 2024). Conventional measures rarely achieve this discrimination, which is central to accurate TRIAD interpretation. Additionally, a single molecular assay can uncover many bioindicators at once, making these multifunctional methods economical for higher-tiered TRIAD assessment. Omic endpoints also trace contaminant effects across trophic levels, revealing mechanisms that propagate through interconnected organisms, such as restructured plant-microbe symbioses (Gonzalez et al., 2018), earthworm gut-to-soil degradation pathways (Wei et al., 2024), and disrupted nutrient exchanges within the soil microbiome (Liu, Y. et al., 2023), all ERA-valuable data that conventional microbial assays overlook. Applied at multiple timepoints, these methods can separate natural background dynamics from contaminant-driven change and track whether that change is transient or persistent, supplying detailed temporal evidence the ecological LoE presently lacks (Jacquiod et al., 2018). The discovery of novel, contaminant-specific biomarkers is facilitated by omics, expanding the library of validated and candidate molecular toxicity indicators (Romdhane et al., 2019). These analyses also detect candidate degrader species, genes, and consortia, informing innovative remediation design (Jiménez-Volkerink et al., 2024), a stated goal of higher-tiered TRIAD and the EU Soil Thematic Strategy.
An established toolkit of multivariate and statistical methods already exists to interpret omic data and convert it into integrable risk biomarkers applicable to TRIAD. Diversity metrics, ordination and clustering techniques, and co-occurrence and causal network analyses correlate taxonomic and functional shifts, or altered biochemical pathways, to measured contaminant and soil physicochemical data, extracting response patterns that can be used to categorize ecological risk (Karimi et al., 2017; Han et al., 2021; van Eijnatten et al., 2025). Dedicated software can predict a community's functional-gene content from taxonomic data alone (Langille et al., 2013), broadening the functional insight available from targeted sequencing procedures. Other tools combine multiple omic datasets from the same samples to find the genes, taxa, and metabolites which change together under contamination, elucidating potential biochemical toxicity mechanisms (Rohart et al., 2017; Singh et al., 2019). Machine learning algorithms have been increasingly employed to process large molecular datasets for diverse field conditions, showcasing their computational power by detecting subtle patterns, and allowing characterization of a vast array of candidate bioindicators. ML models applied to omics data are examined in Part I (Djomehri et al., 2026). Across these analyses, pre-integrating soil physicochemical and contaminant data (chemical LoE) with omic data (ecological or ecotoxicological LoE) correlates molecular responses with contamination levels and site conditions, revealing relationships that inform biomarker identification and sensitivity, parameter weighting, and risk classification for TRIAD ERA.
Linking molecular responses to contaminant concentrations and soil physicochemical data identifies the contaminant levels at which biological harm begins. This insight can refine toxicity threshold values underlying the chemical and ecotoxicological LoEs for different soil conditions (Veerasamy et al., 2023; Romdhane et al., 2019). Such calibration would mark a significant advance for a modernized TRIAD framework. The biomarkers and response patterns generated by omic analyses can be converted into normalized, weighted hazard values ready for aggregation into the IR calculation, as demonstrated by the Transcriptomic Hazard Index developed by Cecchetto et al. (2023). Both multivariate analyses and machine learning show clear potential to build standardized risk scoring protocols using omics, and integrate endpoint data within and between TRIAD LoEs, applicable to diverse field conditions (Campa et al., 2022; Chakraborty et al., 2024; Chen, S. et al., 2024).
Realizing the advantages of incorporating omic methods into a modernized TRIAD requires confronting procedural challenges and an initial investment of labor and expertise. Omic procedures depend on specialized equipment and trained personnel and remain vulnerable to protocol variation that undermines reproducibility, while bioinformatic, multivariate, and machine-learning analyses introduce their own variability through differing pipelines, parameters, and reference databases (Vestergaard et al., 2017). Ongoing advances in sequencing technology continue to lower costs and improve reproducibility, while increasingly user-friendly analysis software, standardized protocols, and comprehensive environmental bioinformatics platforms steadily improve accessibility (Bouchez et al., 2016; Bünemann et al., 2018; Djemiel et al., 2022). The investment is essential for maintaining the relevance of the TRIAD framework. Once established, standardized protocols and advanced analysis tools can be widely adapted to a range of field scenarios, offering researchers, regulators, and stakeholders a practical means to streamline the cost, time, and labor of subsequent assessments.
Despite their growing use in contaminant research, omic methods have rarely been applied within the TRIAD protocol (Gutiérrez et al., 2015; Grassi et al., 2022). This work, Part II, reviews and synthesizes contamination studies assessing molecular omic endpoints, aiming to explore their sensitivity, discrimination, and multifunctional potential as TRIAD indicators, while discussing practical challenges associated with their implementation. Together with the decision science and analytical approaches developed in Part I (Djomehri et al., 2026), the modernized TRIAD framework utilizes omic methods to enhance the resolution of ERA and address the challenges posed by the complexity of contemporary contamination. These additions have the potential to transform TRIAD into a state-of-the-art tool applicable at a global scale, advancing both ERA accuracy and remediation management.
2. The potential of -omic methods for TRIAD ecological risk assessment
Over the past 15 years, high-throughput sequencing has reshaped environmental contamination research (Bouchez et al., 2016; Djemiel et al., 2022). A growing number of studies correlate genomic and transcriptomic indicators with soil toxicity (Karimi et al., 2017; Thakur et al., 2023; Kong et al., 2024). As sequencing technology advances and bioinformatic databases expand, both data generation and interpretation become faster and more cost-effective. However, methodological variation in DNA/RNA extraction and gene annotation undermines the reproducibility and comparability of results (Mocali and Benedetti, 2010; Vestergaard et al., 2017). Developing standardized ISO/OECD protocols for sequencing and analysis, together with methods for tracing procedural uncertainty, is an essential step towards incorporating omic indicators into the TRIAD framework.
Table 1 catalogs the studies reviewed in the following sections by omic method, their study systems, measured endpoints, key findings, and relevance to the TRIAD Lines of Evidence and tiers.
Table 1. -Omic methods applied in soil contamination studies and their relevance to the modernized TRIAD framework.
| Reference | -Omic method | Contaminant/system | Measured endpoints | Key finding | TRIAD relevance (LoE · tier · strength) |
|---|
| TARGETED SEQUENCING — Metabarcoding (taxonomic markers)
|
|
Taraboletti et al. (2023) | 16S metabarcoding | Variably contaminated riverbank soils (agricultural, organochlorine-pesticide, urban) | Bacterial community shifts; core/indicator families | Indicator taxa matched known pollution profiles (agricultural, organochlorine, urban) | Ecological · T2 · Biomarker |
|
Veerasamy et al. (2023) | 16S metabarcoding + PCA/Soil Quality Index | Petroleum-, sludge-, and leakage-contaminated soils | Diversity indices (richness); degrader genera; physicochemical SQI | Lower richness in heavily contaminated soils; SQI combines chemical + ecological LoE data | Ecological + Chemical · T2 · Integration |
|
Ma et al. (2024) | Metabarcoding | Manganese mine sites; moss rhizobial communities | Taxa–metal correlations | Metal-specific taxa correlations; several taxa double as biomarkers and remediators | Ecological · T2 · Discrimination/Biomarker |
|
Al-Bayati et al. (2024) | Metabarcoding | Oil-refinery soils (Pb, Cd) vs urban soils (Cd, Cr, Ni) | Alpha diversity | Lower alpha diversity in heavily contaminated soil distinguished contamination scenarios | Ecological · T2 · Discrimination |
|
Silva et al. (2024) | Metabarcoding | Heavy-metal-contaminated soil under varying abiotic conditions | Community structure/function; metal bioavailability | Abiotic shifts altered community structure/function via metal bioavailability | Ecological · T2 · Discrimination |
|
Wang, J. et al. (2020) | Metabarcoding (90-day microcosm) | Microplastic-amended vs untreated soil | Community succession rate over time | Succession 3x faster under microplastics; turnover rate as a temporal endpoint | Ecotoxicological · T2 · Temporal |
| MICROARRAYS — Functional-gene arrays (GeoChip™) and network analysis
|
|
Somenahally et al. (2021) | GeoChip™ functional array | Arsenic-contaminated rice fields (flooding regimes) | Functional potential for As-reduction/As-methylation
| Flooding regime set As-transformation potential and bioavailable As; management lever | Ecological · T2 · Biomarker/Remediation |
|
Liang et al. (2016) | GeoChip™ + co-occurrence networks | Oil-contaminated soils (gradient) | Functional-gene network connectivity/clustering | Networks reorganized around hydrocarbon-degradation hubs; connectivity fell with oil | Ecological · T2 · Discrimination/Biomarker |
|
Yang et al. (2018) | GeoChip™ + networks | Oil-contaminated soil | C-degradation, N-cycling, methanogenesis genes; greenhouse-gas fluxes | Functional-gene shifts tracked measurable CO2, N2O, and CH4 fluxes | Ecological · T2 · Biomarker |
|
Yin et al. (2015) | GeoChip™ + 16S | Metal-contaminated river sediment (gradient) | Metal-homeostasis/C-metabolism genes; network topology | Metal-resistance genes rose; network gained complexity (vs oil), aiding contaminant discrimination | Ecological · T2 · Discrimination |
| RNA-BASED METHODS — Targeted arrays and whole-transcriptome indexing
|
|
Addis et al. (2022) | RNA-targeted GeoChip™ (cDNA hybridization) | Spartina alterniflora endophytes; weathered crude oil (mesocosm) | Endophytic functional-gene expression | Oil shifted metal-detoxification and hydrocarbon-metabolism gene expression in host endophytes | Ecotoxicological · T2 · Sensitivity |
|
Cecchetto et al. (2023) | Transcriptomic Hazard Index (THI); whole-transcriptome RNA-seq + GSEA | Mollusk, fish, amphibian species (proof of concept) | Normalized hazard score from gene-set change × pathway relevance; 5 hazard classes | THI matched expert judgment (0.96); converts transcriptomic data into a weighted IR input | All LoEs · T2 · Integration/quantification |
| UNTARGETED SEQUENCING — Metagenomics & Metatranscriptomics
|
|
Yan et al. (2016) | Metagenomics + causal network | Uranium-contaminated soils | Functional potential; gene–pathway network | Membrane-transport and carbohydrate genes enriched under U; remediation candidates flagged | Ecological · T3 · Biomarker/Remediation |
|
Yuan et al. (2024) | 16S + metagenomics ('ecological networks') | Soils over a uranium tailings dam | Functional-gene abundance vs geochemical parameters | N, S, Fe, P cycling genes correlated with U-labile fractions | Ecological + Chemical · T3 · Integration/Biomarker |
|
Shi et al. (2023) | Metagenomics | Heavy-metal mining soils; fertilizer regimes | N-cycling gene abundance (napA, norC, nirK) | High-input fertilizer raised denitrification genes, signaling N-loss and N2O risk | Ecological · T3 · Biomarker |
|
Zhao, W. et al. (2023) | Metagenomics | Cr stress; Iris tectorum ± arbuscular mycorrhizal fungi | Resistance genes (chrA, arsB), N/P-cycling genes, bioavailable Cr, rhizosphere network | AMF raised resistance/nutrient genes and lowered bioavailable Cr, protecting the plant | Ecological · T3 · Remediation/Biomarker |
|
Kong et al. (2024) | Metagenomics + co-occurrence networks | Oil-contaminated soil | Hydrocarbon-oxidation ↔ N-cycling coupling; N speciation | Hydrocarbon oxidation coupled to N-cycling, shifting bioavailable N speciation | Ecological · T3 · Discrimination/Biomarker |
|
Liu, J. et al. (2022) | Metagenomics | Petrochemical landfill (5 sites) | PAH-degradation genes; degrader genera | 10 PAH-degradation genes and degrader genera tracked contaminant concentration | Ecological · T3 · Biomarker |
|
Liu, Q. et al. (2024) | Metagenomics + redundancy analysis (RDA) | Three oilfields across a climate gradient | Partitioning soil properties vs petroleum contamination | RDA separated soil-property from contamination effects; aridity inhibited degraders | Ecological + Chemical · T3 · Discrimination/Integration |
|
Jacquiod et al. (2018) | Metatranscriptomics (3 timepoints/yr) | Long-term copper-exposed soil plots | Seasonal expression of microbial functions | Seasonal expression response collapsed under Cu ('decoupling'), scaled to bioavailable Cu | Ecological · T3 · Temporal/Sensitivity |
|
Cabral et al. (2016) | Metatranscriptomics | Contaminated vs pristine mangrove sediments | Heavy-metal-resistance transcripts (CzcA, CusA) | Metal-efflux transcripts (CzcA, CusA) tracked the specific metals present (active exposure) | Ecological · T3 · Biomarker/Discrimination |
|
Gonzalez et al. (2018) | Multi-organism transcriptomics (metatranscriptomics) | Willow roots + microbiome; petroleum-hydrocarbon soil | Plant + fungal + bacterial transcripts (SWEET2, MST1) | Contamination restructured the root-fungi-bacteria system with coordinated transcript shifts | Ecological + Ecotoxicological · T3 · Cross-trophic
|
|
Li, X. et al. (2024) | 16S/18S amplicon + metagenomics | Biodegradable PBAT microplastics; bok choy rhizosphere | Community membership + functional gene content; crop biomass | PBAT impaired rhizosphere carbon metabolism, disrupted biofertilizer benefit, cut biomass | Ecological + Ecotoxicological · T3 · Cross-trophic
|
|
Wei et al. (2024) | Metagenomics | DEHP-spiked soil–earthworm microcosm (42 d) | Degradation efficiency; pathway routing (catechol, protocatechuate) | Earthworms raised DEHP degradation ~65%, with compartment-specific degradation routes | Ecotoxicological · T3 · Cross-trophic/Remediation |
| PROTEOMICS & METABOLOMICS — Standalone
|
|
You et al. (2022) | Proteomics + growth-inhibition assay | Antibiotics (CTC, FLO) + Cu/Cd; Synechocystis | Growth inhibition; proteome; metal-specific responses | Metal-binding antibiotic gave antagonistic toxicity; non-binding gave additive; metal-specific proteome shifts | Ecotoxicological · T3 · Discrimination |
|
Xu, K. et al. (2021) | Metabolomics (LC-MS) | Carbon-based nanomaterials; Eisenia fetida
| Metabolite biomarkers (choline, Phe/Leu ratio) | Sub-lethal biomarkers (choline, Phe/Leu) tied to nanomaterial morphology | Ecotoxicological · T3 · Sensitivity/Biomarker |
|
Shao et al. (2019) | Metabolomics (1H-NMR) | Sodium selenite; Eisenia fetida
| 10 differential metabolic markers; TCA cycle; energy metabolism | Metabolite shifts at the lowest dose, where mortality/growth were non-significant
| Ecotoxicological · T3 · Sensitivity |
|
Van Meter et al. (2018) | Metabolomics | Pesticides (singles + mixtures); green frog liver | Liver metabolites; impaired pathways | 8 pathways impaired by herbicides, 12 by mixtures; separates mixture from single effects | Ecotoxicological · T3 · Discrimination |
|
Wu, X. et al. (2020) | Metabolomics | Polystyrene microplastics; rice (Oryza sativa) | Metabolite profile; carbon/energy metabolism | >70% of metabolites changed; carbon/energy metabolism and TCA cycle depressed | Ecotoxicological · T3 · Sensitivity/Biomarker |
|
O'Hara & Longstaffe (2023) | Metabolomics (1H-NMR) | PFAS (PFOA, PFOS); Arabidopsis thaliana
| Soluble sugars, amino acids, TCA-cycle intermediates | Stress-metabolite accumulation raised TCA activity (opposite direction to rice) | Ecotoxicological · T3 · Discrimination/Biomarker |
| MULTI-OMICS — Integrated
|
|
Li, C. et al. (2023) | Metabolomics + transcriptomics + proteomics | Cd, PAHs, and co-exposure; ryegrass (Lolium perenne) | Linoleic-acid oxidation, lignin biosynthesis; co-exposure-specific metabolites | Co-exposure disrupted linoleic-acid oxidation and lignin biosynthesis; mixture-specific biomarkers | Ecotoxicological · T3 · Discrimination |
|
Zhang, Y. et al. (2024) | Transcriptomics + metabolomics + physiology | Pb stress; Pb-sensitive vs Pb-resistant Hydrangea macrophylla
| Antioxidant enzymes, soluble sugar, membrane integrity; cutin/suberin/wax pathway | Tolerance traced to cutin/suberin/wax biosynthesis; physiology confirmed phenotype | Ecotoxicological · T3 · Discrimination/Biomarker |
|
Tang et al. (2020) | Proteomics + metabolomics | Low-level arsenic-contaminated soil; Eisenia fetida
| Proteome/metabolome; biomarkers (asparagine, dimethylglycine) | Molecular toxicity (osmotic stress, immune dysregulation) with no survival/weight change | Ecotoxicological · T3 · Sensitivity/Biomarker |
|
Zhao, Y. et al. (2024) | Transcriptomics + metabolomics + gut microbiome | Biodegradable (PLA) vs conventional (PVC, LDPE) microplastics; Eisenia fetida; biogas-slurry irrigation | Gut community; host stress responses | Polymer-dependent gut shift mirrored host responses (PLA vs PVC/LDPE) | Ecotoxicological + Ecological · T3 · Discrimination |
|
Boughattas et al. (2024) | 16S + metabolomics (DIABLO/mixOmics) | 2,4-D, microplastics, and combination; Eisenia andrei
| Metabolite–taxa correlations across treatments | Co-exposure most pronounced; metabolites of oxidative stress, energy, nucleic-acid metabolism | Ecotoxicological · T3 · Discrimination/Integration |
|
Romdhane et al. (2019) | 16S + metabolomics (DIABLO/mixOmics) | Two β-triketone herbicides (leptospermone, sulcotrione); soil community + metabolome | OTU–metabolite signatures by dose | Distinct OTU-metabolite signature per herbicide; herbicide-specific biomarkers | Ecological · T3 · Discrimination/Biomarker |
|
Li, X. et al. (2019) | Metabolomics + PICRUSt + enzyme assays | Long-term PAH soil + fresh PAH input | Soil enzymes (dehydrogenase, urease); 43 metabolites; predicted pathways | Fresh PAH inhibited soil enzymes; 43 metabolites changed across key pathways | Ecological + Ecotoxicological · T3 · Biomarker/Integration |
|
Wu, C. et al. (2022) | 16S + metabolomics (LEfSe, PICRUSt) | Plastic-mulch-contaminated soil | Distinctive taxa; predicted degradation pathways | Clostridium_sensu_stricto_1 correlated with caprolactam degradation; remediator discovery | Ecological · T3 · Biomarker/Remediation |
|
Dong et al. (2024) | Integrated multi-omics
| Plastic residues (PLA vs PE), 120 d; farmland, woodland, wetland soils | Community composition, C/N/P-cycling taxa, metabolites; enzymes, nutrients, pH | Soil-type-specific shifts; PLA disturbed enzymes/nutrients more than PE | Ecological · T3 · Discrimination |
|
Han et al. (2024);
Han et al. (2021) | Metagenomics + root-exudate metabolomics; PICRUSt | PBAT (biodegradable) vs PE microplastics; pakchoi; multiple soil types | C/N/P/S-cycling potential; plant growth; degrader genera; causal model | PBAT (not PE) suppressed cycling and growth dose-dependently; causal model explained 87% of variation | Ecological + Ecotoxicological · T3 · Discrimination/Cross-trophic
|
|
Liu, Y. et al. (2023) | Metagenomics + metabolomics | Polypropylene vs rubber-crumb microplastics; peanut rhizosphere | N-cycling genes; metabolites; root-cell damage, oxidative stress | PP and rubber crumb disrupted N-cycling differently; rubber crumb more severe | Ecological + Ecotoxicological · T3 · Discrimination |
|
Jiménez-Volkerink et al. (2024) | Metagenomics + metabolomics (genome-resolved) | Oxy-PAH (BaAQ) from creosote-contaminated soil; degrading consortium | Degradation pathway distribution; nutritional interdependencies; horizontal gene transfer | Degradation split across consortium via division of labor and HGT; informs bioremediation | Ecological · T3 · Remediation/Mechanism |
2.1 Targeted high-throughput sequencing and microarrays
Targeted DNA sequencing, or metabarcoding, uses primers to select specific taxonomic (16S rRNA for bacteria/archaea, 18S rRNA for eukaryotes, ITS for fungi) or functional genes for Polymerase Chain Reaction (PCR) amplification and sequencing. For taxonomic markers, this approach generates Operational Taxonomic Units (OTUs) resolvable from phylum to species levels, along with their relative abundances, facilitating the tracking of community shifts through diversity metrics such as alpha and beta diversity or species’ evenness and richness. Because taxonomic markers are highly conserved at the regions where primers anneal, a single primer pair can cover many species. While primers can also target functional genes, the lack of highly conserved binding sites across species limits their coverage. Functional-gene amplicon sequencing remains uncommon in community-level contamination assessment, unless coupled with qPCR methods that quantify gene copy number (Penton et al., 2013; Lan et al., 2024).
For functional gene markers, high-density microarrays offer a more practical route, using over 150,000 pre-designed probes to survey many functional genes and their variants at once. Shifts in the presence and relative abundance of genes indicate changes in the community's functional potential for processes such as growth, reproduction, nutrient cycling, contaminant degradation, and stress response. Unlike primer-based methods, microarrays use probes that report abundance through a fluorescent hybridization signal that scales with how much target sequence binds to each probe. Because the targets are preset, no later sequencing is required. Microarrays are closed format technologies, unable to detect unknown diversity.
Co-occurrence network analysis, an emerging approach in soil health research, offers greater sensitivity by detecting potential interactions among taxa or genes, not only changes in their abundance or diversity. Applicable to taxonomic or functional datasets, this method constructs networks with nodes representing taxa or genes and edges reflecting correlations in their abundances across samples. These networks identify clusters of taxa or genes that respond similarly to environmental conditions. Incorporating soil physicochemical and contaminant data further uncovers and distinguishes associations between environmental drivers, contaminants, and community structure (Karimi et al., 2017; van Eijnatten et al., 2025). For TRIAD implementation, targeted sequencing and analysis methods are increasingly affordable and provide multiple biomarkers from a single procedure, making them well-suited for tier 2 assessments.
Changes in soil community composition correlate with contamination levels, providing sensitive, indicators for TRIAD. Taraboletti et al. (2023) recorded bacterial community shifts for variably-contaminated riverbank soils. Core families shared among sites matched reference taxa known to indicate agricultural and organochlorine-pesticide pollution (Burkholderiaceae, Nitrosomonadaceae, Pedosphaeraceae), while Xanthobacteraceae and Methyloligellaceae aligned with urban, high-carbon conditions. As pollutant-associated taxa databases expand, this approach offers an increasingly affordable route to TRIAD-ready biomarkers. Linking diversity indices to organic contamination, Veerasamy et al. (2023) documented lower bacterial richness in heavily contaminated sludge and leakage soils than in aged petroleum-exposed soil, with hydrocarbon-tolerant and degrading genera such as Paracoccus and Pseudomonas present in all samples. The researchers also built a Soil Quality Index from physicochemical parameters using Principal Components Analysis (PCA) and a minimum dataset, an approach that could be used to combine chemical LoE data with molecular community data (ecological LoE) in TRIAD assessments. At manganese mine sites, moss rhizobial communities varied with measured metal concentrations, with taxa such as Planctomycetes correlated positively with Mn and Cd, Acidobacteria with Ni, and Firmicutes associated negatively with Pb (Ma et al., 2024). Cyanobacteria and Actinomycetes showed strong tolerance to mixed-metal contamination. Several metal-associated taxa emerged as both contaminant-specific biomarkers and candidate remediators, highlighting the sensitivity and discriminatory power of metabarcoding for deriving toxicity indicators applicable to TRIAD.
Metabarcoding for taxonomic markers distinguishes how microbial communities respond to co-occurring contaminants, varying abiotic conditions, and temporal exposure. Comparing oil refinery soils enriched with Pb and Cd against urban soils high in Cd with moderate Cr and Ni, Al-Bayati et al. (2024) found lower alpha diversity in the more heavily contaminated refinery community, distinguishing the two contamination scenarios and illustrating how microbial diversity indices can supply sensitive parameters for the ecological LoE. Short-term shifts in temperature, moisture, and UV radiation significantly altered the structure and function of a heavy-metal-contaminated soil community, partly by changing the bioavailability of the metals themselves (Silva et al., 2024). These results show how abiotic conditions modulate contamination response, underscoring that climate variables must be accounted for when interpreting contaminant effects in TRIAD. In a 90-day laboratory microcosm, bacterial community succession progressed three times faster in microplastic-amended soil than in untreated soil, with species’ composition diverging further from initial conditions at each timepoint (Wang, J. et al., 2020). A single-timepoint test would miss this increased community turnover rate, which sequencing captures as an accessible, time-resolved endpoint for the ecotoxicological LoE.
Despite their broad coverage, primer-based taxonomic sequencing remains limited by the sheer diversity of soil microbes. The method can detect unknown OTUs that represent novel diversity, however, lineages whose binding sites diverge too far from the primer sequence are never amplified and remain invisible, leaving gaps in the community profile. To address this challenge, researchers continually develop and refine primers to be more universally applicable, (Tedersoo et al., 2016) and also to mitigate PCR amplification bias arising from primer-template mismatch (Bergkemper et al., 2016). To broaden coverage across functional sequence variants, microarrays use several probes to capture different variants of each gene, grouped by gene family. Additionally, each probe binds any sequence that matches by 90% or more, capturing several closely-related variants. Although microarrays avoid primer-specific amplification bias, which is systematic and correctable, if more starting material is needed to enable gene/transcript detection, an initial whole-community step (for DNA or RNA) amplifies all sequences in the sample. This procedure introduces the stochastic distortions unavoidable with any amplification process (Vestergaard et al., 2017). Due to the exponential nature of sequence replication, amplification drift over-represents whichever sequences are copied first by chance. Rare community members may also under-amplify or drop out, and erroneously replicated strands add noise. Advances in amplification-free methods (Zhou et al., 2015), alongside improvements to experimental design and analytical methods (Mocali and Benedetti, 2010; Vestergaard et al., 2017), continue to reduce these inaccuracies and support standardization, broadening their practicability for contaminant-based assessments such as TRIAD.
Microarrays such as GeoChip™ (Shi, Z. et al., 2019) target genes involved in nutrient cycling, stress response, metal resistance, and pollutant degradation, generating large datasets from which researchers can identify numerous toxicity indicators through functional diversity and co-occurrence network analyses. Somenahally et al. (2021) linked continuously-flooded, arsenic-contaminated rice fields to a higher functional potential for microbial As-reduction and As-methylation, which accounted for increased bioavailable As (III) and methyl-As in soil pore water and rice grain. Intermittently flooded fields showed reduction in both these As-transforming processes and overall As levels in rice, pinpointing functional gene biomarkers of human health risk and a practical management intervention. In oil-contaminated soils, GeoChip™-derived functional gene co-occurrence networks exhibited reduced connectivity and weaker clustering, reorganizing around alkane- and PAH-degradation genes as their most connected hubs (Liang et al., 2016). Gene connectivity fell as oil concentration rose, linking the contamination gradient directly to network simplification. Negative interactions prevailed among the degradation genes, indicating potential competition among the species that carry them. This restructuring marks a clear network-level signature of community adaptation to hydrocarbon contamination, capturing interaction details abundance metrics alone would overlook.
At another oil-contaminated site, GeoChip™-derived networks also exhibited simplification, with reduced nodes and connectivity. Declines in C-degradation and N-cycling genes correlated with reduced CO2 and N2O fluxes, while methane-production genes increased, linked to the elevated CH4 emissions observed (Yang et al., 2018). These coordinated shifts tie functional gene patterns in contaminated soil directly to measurable greenhouse-gas emissions. Combining GeoChip™ functional data with 16S taxonomic profiling in metal-contaminated river sediment, Yin et al. (2015) found higher abundances of metal-homeostasis and C-metabolism genes correlated with metal-resistant species across the contamination gradient. In contrast to the oil-contaminated networks described, network interactions here displayed increased complexity, with metal-resistant populations becoming more interconnected. Thus, network topology may have the potential to help discriminate metal from hydrocarbon contamination. Microarray studies deliver sensitive functional biomarkers and network-level signatures that discriminate contaminant types, generating the large datasets needed to develop predictive ML models for TRIAD.
Targeted RNA-seq provides gene expression snapshots indicating differentially expressed genes (DEGs), which signal up- or down-regulated functional pathways linked to contamination, and is primarily used for profiling functional transcripts. However, primer-based RNA methods for functional targets face the same coverage issues as DNA-based methods, yielding only a narrow range of sequence variants and select few transcripts per assay, limiting their use in broad-scale contaminant research. The microarray approach can extend to transcripts as well. Reverse-transcribed RNA (cDNA) hybridized to a GeoChip™ measures the expression of thousands of preset functional genes at once. In a greenhouse mesocosm study, Addis et al. (2022) inoculated seedlings of the dominant marsh grass Spartina alterniflora with microbial communities from oiled and unoiled Louisiana salt-marshes, applied weathered crude oil, and measured the resulting endophytic transcript expression by hybridizing reverse-transcribed RNA (cDNA) to a GeoChip™ array. Oil exposure up-regulated metal-detoxification genes, apart from chromium, and shifted the expression of hydrocarbon-metabolism,
C, P, and secondary-metabolism genes, changes with possible consequences for host plant fitness. Measuring expressed genes captured the endophytic community's realized response to contamination within the host, a sensitive endpoint well suited to the ecotoxicological LoE.
Notably, a meta-analysis by Rocca et al. (2015) found only a weak positive correlation between gene abundance and corresponding processes, and no significant correlation for transcript abundance, attributable to rapid RNA degradation in the environment and the complex relationship between transcript levels and enzyme activity. Yet the same rapid turnover that weakens this correlation, makes transcript abundance a sensitive, real-time indicator of dynamic stress response, registering rapid up- or down- regulation that single timepoint or before-and-after sampling would miss. As extraction and sequencing methods improve, transcriptomics offers temporal resolution of stress-response and contaminant-degradation pathways valuable for the ecological LoE.
2.2 Untargeted sequencing
Untargeted sequencing, though more resource-intensive, sequences all extracted DNA/RNA from an environmental sample, enabling both phylogenetic and functional bioinformatic analysis without the specific primer biases of targeted approaches. Metagenomics captures the full scope of community composition and functional gene content, while metatranscriptomics detects expressed genes (Djemiel et al., 2022). Within organisms, whole-genome profiling establishes functional potential, including the presence of contaminant resistance or degrader genes. Transcriptomic methods identify which genes are activated or disrupted under contamination. Genomics/metagenomics can pinpoint remediation potential within a community or organism, and transcriptomics/metatranscriptomics can verify that potential, a much-needed capability for TRIAD assessment in the face of modern pollution.
Because no genes are preselected, ‘global’ sequencing can also reveal complex patterns in contaminant responses spanning community, population, and organism levels, that remain undetectable through targeted approaches. A single metagenomic analysis delivers both taxonomic and functional signals at once, and can yield numerous novel bioindicators, offering comprehensive data suited for higher-tiered TRIAD assessments. These data also expand environmental reference databases, both for improved gene annotation (Djemiel et al., 2022), and pollutant-indicator matching, as in Taraboletti et al. (2023). Conventional global sequencing includes a PCR step which introduces amplification biases such as uneven amplification of GC-rich or GC-poor fragments, and duplicate reads. Amplification-free methods omit this step, avoiding these biases. Recent work applying amplicon-free, long read sequencing of DNA in metal-contaminated soils resolved community response at the strain level, illustrating their value for tier 3 assessments as the technology becomes more accessible (Isgandarov et al., 2026).
Figure 3 summarizes the sequencing-based omic methods covered here, from targeted DNA and RNA approaches to global sequencing. Additionally, untargeted omic datasets are essential for training ML models (Campa et al. 2022; Chakraborty et al., 2024) that can inform and streamline lower-tiered protocols, examined further in Part I (Djomehri et al., 2026).
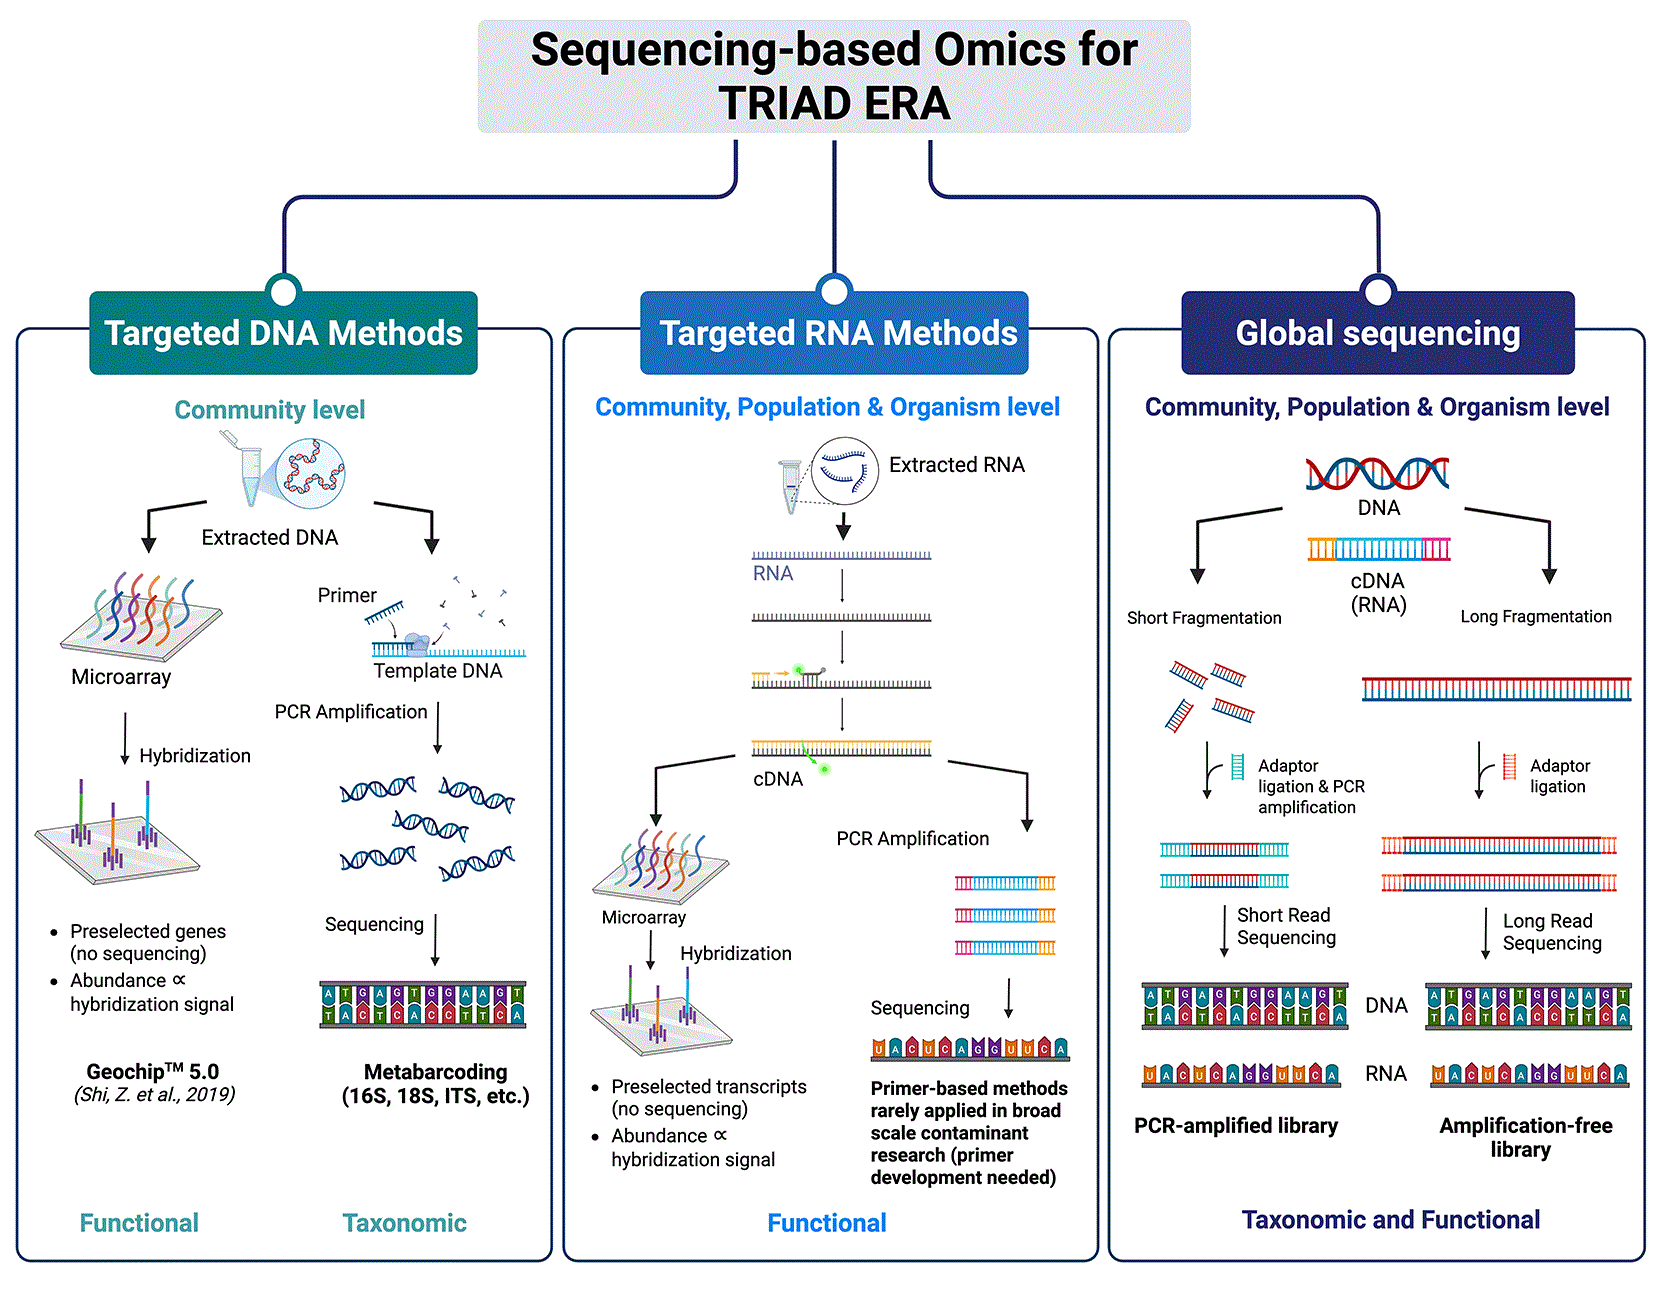
Figure 3. Sequencing-based omic methods proposed for TRIAD ecological risk assessment.
Targeted DNA methods select preset genes: microarrays (e.g., GeoChip™ 5.0) identify and quantify functional genes without sequencing; metabarcoding amplifies and sequences taxonomic markers (16S, 18S, ITS). Targeted RNA approaches reverse-transcribe RNA to cDNA, then measure functional gene expression by microarray hybridization or primer-based methods: narrow variant coverage has limited the latter in broad-scale contaminant research. Global sequencing probes all DNA, or RNA (via cDNA) within a sample. Established protocols create short nucleic acid fragments that are adaptor-ligated and PCR-amplified for short-read sequencing; emerging long read technologies are adaptor-ligated and proceed without amplification. Amplification-free methods are a developing field with promise for improving accuracy in ERA. Created with BioRender.
Global sequencing delivers detailed discoveries for high-risk sites, underscoring the value of these methods for TRIAD assessments. In uranium-contaminated soils, Yan et al. (2016) found significantly altered microbial communities and metabolic functional potential, with membrane transport and carbohydrate metabolism genes enriched in contaminated soil, functions considered essential to the community under U stress. Actinobacteria and Proteobacteria emerged as dominant phyla, candidates for U-contaminated soil remediation. In a causal network linking functional gene groups to metabolic pathways, membrane transport and carbon metabolism genes were among the most connected, both consistent with a central role for membrane transport in microbial response to U toxicity. Combining 16S and metagenomic analysis of soils covering a uranium tailings dam, Yuan et al. (2024) constructed 'ecological networks' linking functional gene abundances to geochemical parameters, documenting specific N, S, Fe, and P cycling genes that strongly correlated with U-labile fractions. Together, the methods used in these studies surface novel biomarkers and advance candidate species for remediation, making them applicable to tier 3 TRIAD protocols.
Agricultural risk assessments frequently utilize untargeted sequencing to understand the impact of contamination on plant health and soil function. In HM-contaminated mining soils, N-cycling microbial gene abundance varied with fertilizer type, dose, and frequency, with high-input regimes sharply increasing nitrogen-reduction genes (napA) and denitrification genes (norC and nirK) (Shi et al., 2023). This elevated denitrification potential can signal a risk of gaseous nitrogen loss, limiting nutrient availability, and potentially increasing N2O emissions, establishing functional markers with direct implications for fertilization management of contaminated soils. Under Cr stress, Iris tectorum inoculated with arbuscular mycorrhizal fungi (AMF) retained greater biomass and photosynthetic pigment content than uninoculated plants. Metagenomics traced this protection to the rhizosphere community. AMF increased the abundance of heavy-metal-resistance genes (chrA, arsB) and N/P-cycling genes, lowered bioavailable Cr levels, and enhanced the complexity and stability of the rhizosphere network (Zhao, W. et al., 2023). The functional gene resolution enabled by untargeted metagenomics can both flag contamination risks and the mechanisms behind bioremedial interventions, advancing ERA and management strategies for agricultural soils.
Untargeted sequencing uncovers hidden contaminant-response mechanisms, reveals sensitive biomarkers, and separates environmental from contamination effects, capabilities central to advanced TRIAD ERA. Kong et al. (2024) found that oil contamination shifted microbial communities toward petroleum-hydrocarbon (PH)-degrading populations, with co-occurrence network analysis coupling hydrocarbon oxidation to the nitrogen-cycling processes of dissimilatory nitrate reduction to ammonium (DNRA), N-fixation, and N-immobilization. This coupling shifted N speciation, raising ammonium while lowering nitrate and nitrite, altering the forms of bioavailable nitrogen available and disrupting ecosystem N cycling. The shift toward hydrocarbon-degrading populations recurs in petroleum-contaminated environments (Cabral et al., 2022). Sampling five sites at a petrochemical landfill, Liu, J. et al. (2022) detected 10 PAH-degradation-related genes whose abundances were linked to concentration, alongside enriched degrader genera such as Bradyrhizobium, Mycobacterium, and Anaeromyxobacter. Together, these offer sensitive functional and taxonomic biomarkers tying contaminant levels directly to microbial degradation potential, highlighting the multipurpose nature of metagenomic approaches. In three oilfields spanning a climate gradient, redundancy analysis (RDA) separated the effects of soil properties from those of petroleum contamination on the microbial community. Low soil moisture in arid climates was identified as a distinct driver inhibiting the accumulation of petroleum-degrading microbes (Liu, Q. et al., 2024). Untargeted sequencing supplies the high-resolution data that allow environmental and contamination effects to be partitioned, an essential step for accurate TRIAD assessments (Chapman and Hollert, 2006; Volchko et al., 2019).
Untargeted metatranscriptomic sequencing highlights DEGs as powerful biomarkers signaling contaminant presence and its functional consequences. This approach is suited to temporal study designs in ERA, an area which critically needs more focus in TRIAD in order to increase assessment accuracy. Jacquiod et al. (2018) compared soil metatranscriptomes among plots with varying long-term copper exposure, at three timepoints over a year. In the control, 309 microbial functions exhibited seasonal shifts in expression, while in the most contaminated plot this temporal response fell sharply. The community had lost its capacity to react to environmental fluctuations. This functional 'decoupling' was proportional to bioavailable Cu, and would be invisible to single-timepoint sampling, illustrating the benefit of temporal studies incorporating untargeted RNA-seq into TRIAD. The identity of contamination-triggered transcripts provides a snapshot of current exposure to specific contaminants. A metatranscriptome comparison of contaminated and pristine mangrove sediments detected elevated abundances of transcripts encoding heavy metal resistance proteins, among them the cobalt-zinc-cadmium efflux pump CzcA and the copper/silver efflux pump CusA, transporters that expel toxic metal ions from the cell (Cabral et al., 2016). This expression tied directly to the specific metals present, marking active exposure rather than historical contamination.
As a proof of concept for incorporating transcriptomic evidence into TRIAD, Cecchetto et al. (2023) developed a Transcriptomic Hazard Index (THI) which combines the degree of gene-set change under contamination with the biological relevance of the affected pathways, classifying hazard into five levels. Developed and confirmed for mollusk, fish, and amphibian species, the THI reached a 0.96 Spearman correlation with expert judgment, indicating that reproducible, quantitative indexing can calibrate or complement expert assessment of transcriptomic hazard. The normalized output is a weighted LoE endpoint that integrates directly into quantitative Weight-of-Evidence ERA frameworks such as TRIAD. The index draws on transcriptomic data alone, reflecting the premise that molecular data inform risk only once combined with the wider evidence base. Because the THI is assembled from interchangeable components (a gene-set database, enrichment analysis, and pathway weighting), the statistical approach is adaptable to terrestrial organisms and other ecological conditions. The sophisticated omic platforms and bioinformatic tools used throughout these studies to quantify contaminant responses could readily generate indices of this kind, giving molecular endpoints a standardized, weight-able form for TRIAD assessment.
Global sequencing spanning trophic systems captures how contaminant effects cascade through interconnected organisms, providing critical data to enhance tier 3 TRIAD assessments. Plant root exudates can desorb organic pollutants from soil particles, raising their bioavailability. Higher bioavailability heightens potential toxicity to the soil community while also stimulating pollutant-degrading microbes active in contaminant removal, illustrating an interspecies mechanism for remediation (Lü et al., 2024). Gonzalez et al. (2018) sequenced transcripts from willow roots and their microbiome in PH-contaminated soils, discovering that contamination restructured the tree root-fungi-bacteria system. The fungal community shifted from saprotroph-dominated Ascomycota toward an uncharacterized, likely ectomycorrhizal Basidiomycota group that aided nutrient uptake and stress tolerance. The willow upregulated transcripts encoding the sugar-export transporter SWEET2, while the Basidiomycota upregulated the sugar-import transporter MST1, expression consistent with increased carbon transfer. A bacterial rhizosphere partner contributed to hydrocarbon degradation and stress reduction. Only untargeted, multi-organism sequencing could resolve this coordinated response, demonstrating the value of these methods for interpreting risk.
Agricultural studies follow the downstream effects of contamination on crops. Li, X. et al. (2024) combined 16S/18S amplicon sequencing for rapid identification of community membership, and untargeted metagenomics for functional gene content, resolving that biodegradable PBAT microplastics shifted the bok choy (Brassica chinensis) rhizosphere community and impaired carbon-metabolic function. These changes disrupted the beneficial effects of a Bacillus biofertilizer and reduced crop biomass, linking contaminant effects on soil microbes directly to plant growth. Metagenomics can detect cross-trophic effects extending to soil fauna as well. In a soil-earthworm laboratory microcosm spiked with environmentally-relevant doses of di-(2-ethylhexyl) phthalate (DEHP), Wei et al. (2024) found earthworm-amended soil degraded DEHP roughly 65% more efficiently than earthworm-free soil over 42 days. Each compartment (soil, casts, earthworm gut microbiome) associated with a distinct degradation route: the catechol pathway in soil, the protocatechuate pathway in earthworm casts, and both in the earthworm gut. Earthworms raised soil humic content and stimulated DEHP-degrading microbes through gut interactions, spreading the degradation potential throughout the animal-soil system via the casts. This study exemplifies an advanced, molecular tier-3 ecotoxicological assessment, sensitive enough to expose fine mechanistic detail and point toward valuable remediation pathways. Integrating multi-omic layers can illuminate these multi-trophic contaminant-triggered interactions further, connecting genetic potential to the proteins and metabolites produced.
2.3 Proteomics, metabolomics, and multi-omics
Proteomic and metabolomic analyses indicate contamination-activated biochemical pathways through differentially expressed proteins (DEPs) and differentially abundant metabolites (DAMs). While powerful as standalone methods, their integration with genomic and transcriptomic data connects biochemical response to underlying gene expression, providing a more comprehensive molecular picture of contaminant risk. By further integrating multi-omics endpoints with soil physicochemical, ecotoxicological, and ecological data, contaminant researchers have resolved sensitive response patterns that can feed directly into TRIAD Line of Evidence weighting and integrated risk (IR) calculation.
Extraction challenges such as sample heterogeneity and low extraction yield remain limitations of environmental proteomics (Bouchez et al., 2016; Djemiel et al., 2022), though ongoing protocol improvements enhance reliability, such as optimized extraction methods that increase protein recovery from soil (Herruzo-Ruiz et al., 2021). Stand-alone proteomics resolves the molecular mechanisms behind contaminant toxicity, including responses invisible to standard toxicity endpoints. Pairing a growth-inhibition assay with proteomics illustrates how this molecular data can enhance the ecotoxicological LoE. You et al. (2022) tested two antibiotics, together with Cu and Cd, against Synechocystis (cyanobacteria) cultures, tracing their combined toxicity to whether the antibiotic bound the metals. Chlortetracycline complexed Cu and Cd, an interaction that reduced growth inhibition and lessened disruption of the proteome. Florfenicol did not bind the metals, so the resulting toxicity was additive, lowering ATP synthesis, photosynthesis, and carbon fixation. Proteomics also separated metal-specific responses, with Cu increasing iron-sulfur assembly and Cd disrupting cyclic electron transport. These high-resolution co-contaminant distinctions are discernible only through molecular-level assays.
Stand-alone metabolomics elucidates active functional pathways across trophic levels with high sensitivity, and has been applied extensively in environmental research (Shahid et al., 2023; Simpson, 2023; Monteiro et al., 2024). The approach detects metabolic disruption at sublethal exposures and identifies the specific pathways affected, two strengths that directly serve higher-tier TRIAD ecotoxicology. In Eisenia fetida exposed to carbon-based nanomaterials, key altered metabolites served as toxicity biomarkers, including choline depletion, indicating membrane damage, and phenylalanine/leucine ratio shifts, suggesting energy metabolism disruption, with these effects tied to nanomaterial morphology (Xu, K. et al., 2021). Another earthworm study found 10 differentially abundant metabolic markers in selenite-exposed Eisenia fetida, disrupting the tricarboxylic acid (TCA) cycle and energy metabolism. Metabolite perturbations appeared at the lowest concentrations tested (10 mg/kg), where mortality and growth inhibition remained non-significant, highlighting the sensitivity of metabolomics for early detection of contaminant effects (Shao et al., 2019). In green frogs (Lithobates clamitans), pesticide exposure altered the abundances of dozens of liver metabolites. Pathway analysis mapped these changes to eight biological pathways impaired by the herbicide treatments and 12 under mixed-pesticides (Van Meter et al., 2018). By tracing altered metabolites to specific pathways and separating mixture effects from those of single compounds, this approach delivers the sensitivity and discrimination that the TRIAD ecotoxicological LoE critically lacks.
Crop studies expose similarly extensive disruptions. Wu, X. et al. (2020) documented significant change in over 70% of metabolites in rice plants (Oryza sativa) grown in polystyrene-contaminated soil. Amino acids, organic acids, and sugars declined as microplastic concentrations increased, affecting carbon and energy metabolism, including the TCA cycle. Under PFAS exposure, Arabidopsis thaliana accumulated soluble sugars, several amino acids, and TCA-cycle intermediates, a metabolic signature the authors interpreted as a stress response that pushed TCA-cycle activity upward (O'Hara and Longstaffe, 2023). The two studies move in opposite directions, microplastics depleting metabolite pools in rice and PFAS elevating them in Arabidopsis, demonstrating that metabolomics discriminates the actual metabolic response of a given organism to a given contaminant. The approach identifies sensitive biomarkers, adding to the growing library of candidate metabolic indicators for ERA, and building the reference knowledge needed to standardize molecular endpoints within TRIAD.
An early study on contaminated soil community metabolomics (Jones et al., 2014) proposed that combining metabolomics with metagenomics provides a 'high-throughput solution', connecting taxonomy with function, an approach essential for building reference databases and predictive ML models for use in TRIAD. Integrating genomic/transcriptomic with proteomic/metabolomic data enables direct comparison of gene presence or expression to the proteins and metabolites actually produced, addressing the weak gene-to-process correlations described by Rocca et al. (2015) and linking molecular biomarkers to specific contaminants and soil conditions for evidence-based TRIAD assessments (Li, X. et al., 2019; Dong et al., 2024).
Figure 4 shows how integrated multi-omics data links the genomic, transcriptomic, proteomic, and metabolomic layers to contamination-response pathways, TRIAD Line of Evidence assignment, and integrated risk calculation.
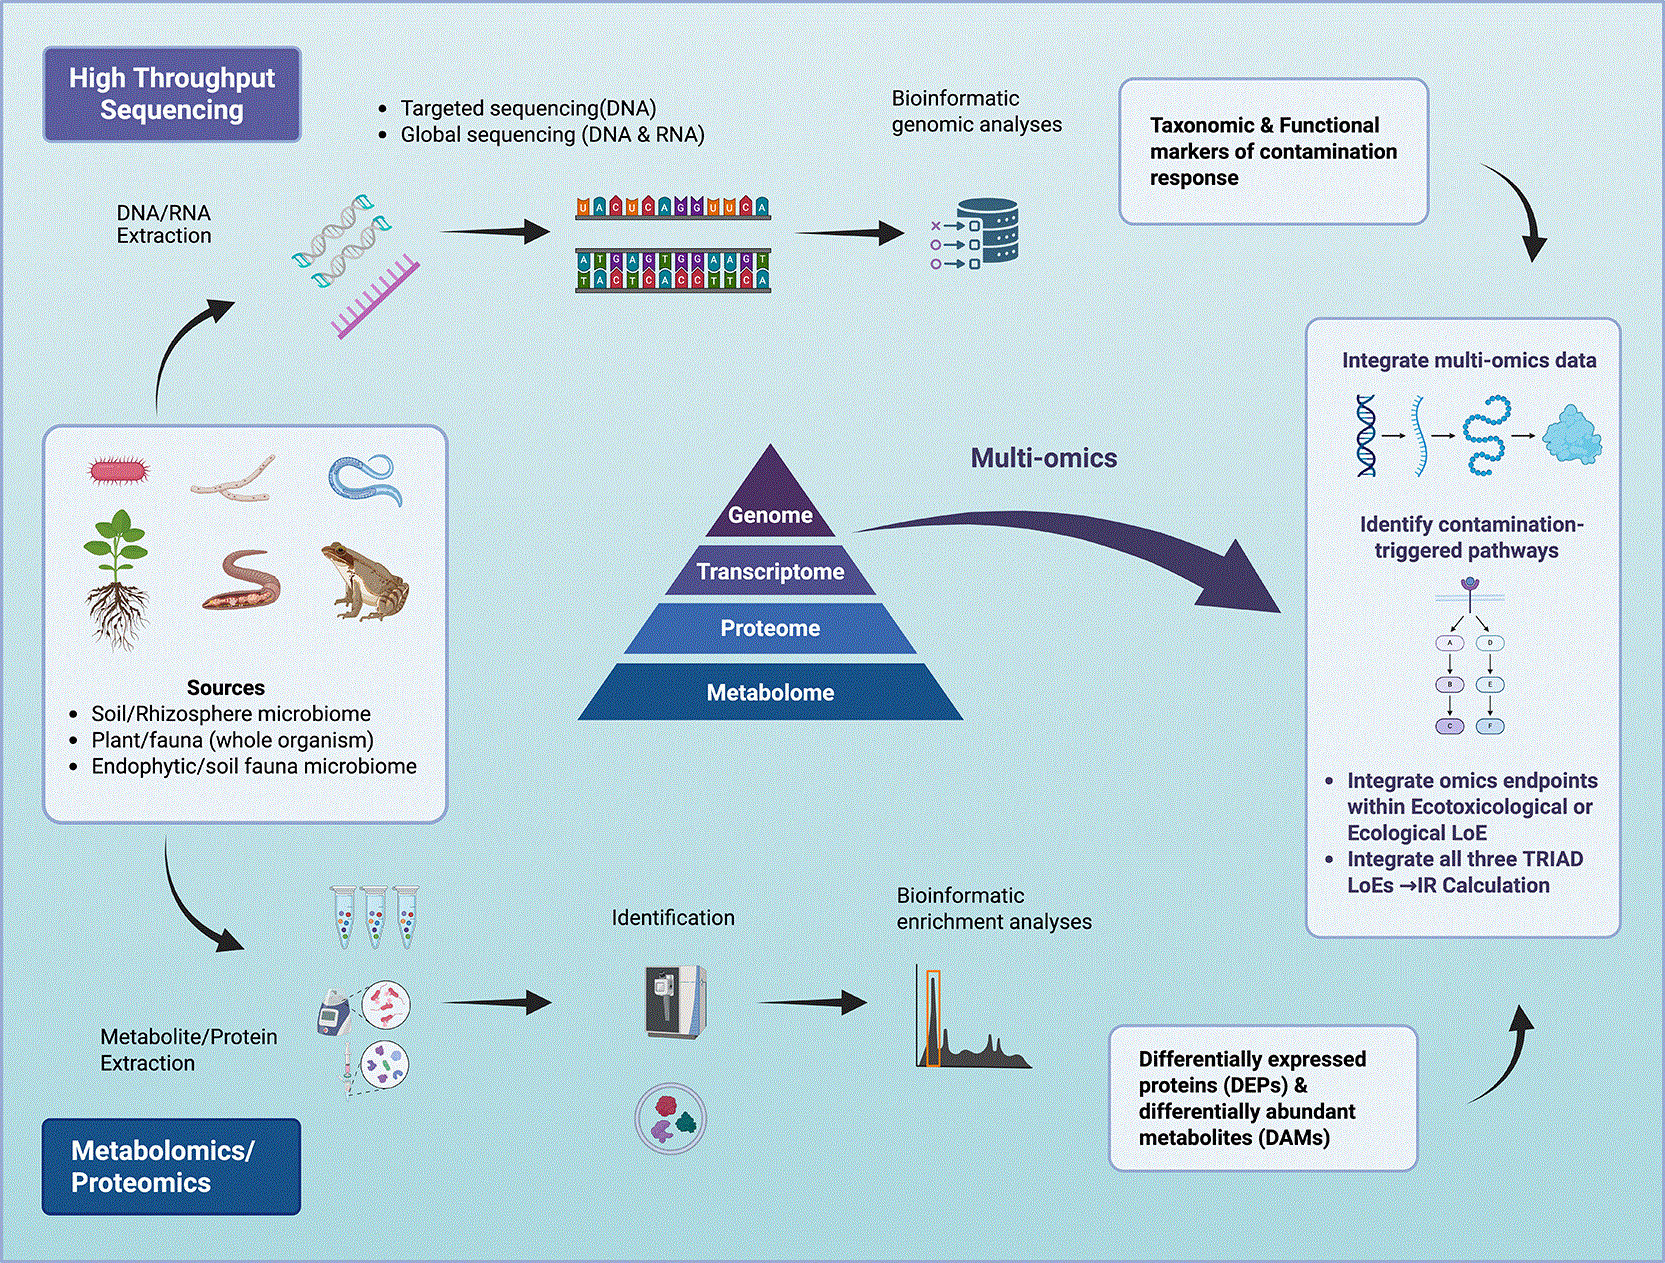
Figure 4. Multi-omics integration for TRIAD ecological risk assessment.
Molecular data from soil/rhizosphere microbiomes, whole plants and fauna, and endophytic or soil-fauna microbiomes are generated by two streams: high-throughput sequencing, producing taxonomic and functional contamination markers, and metabolomics/proteomics, producing differentially expressed proteins (DEPs) and differentially abundant metabolites (DAMs). Integrating the genomic, transcriptomic, proteomic, and metabolomic layers identifies contamination-triggered pathways, whose endpoints are assigned to the ecotoxicological or ecological Lines of Evidence and combined across all three TRIAD LoEs to calculate integrated risk (IR). Created with BioRender.
Combining transcriptomic and metabolomic data from organisms with well-documented genomes connects biochemical responses to gene expression, providing the resolution needed to improve TRIAD's ecotoxicological LoE. Integrating metabolomics, transcriptomics, and proteomics, Li, C. et al. (2023) examined ryegrass (Lolium perenne) under Cd, PAHs, and co-contaminant scenarios. The co-exposure disrupted linoleic acid oxidation, elevating oxidative stress, and inhibited lignin biosynthesis, weakening cell wall defense. Comparison against the single-contaminant treatments isolated biomarkers specific to the combined exposure, demonstrating the discrimination potential of multi-omics. Comparing Pb-sensitive and Pb-resistant Hydrangea macrophylla cultivars, Zhang, Y. et al. (2024) paired transcriptomic and metabolomic profiling with physiological measurements. The resistant cultivar showed higher antioxidant enzyme activity, greater soluble sugar accumulation, and improved root membrane integrity under Pb stress. Conjoint analysis of the transcriptome and metabolome traced tolerance to the cutin, suberin, and wax biosynthesis pathway, with the physiological data confirming the resistance phenotype.
Integrated multi-omics applied to earthworm studies emphasizes the sensitivity of these methods for detecting contaminant effects invisible to conventional assays. Tang et al. (2020) chose multi-omics to address the poorly understood toxicity of low-level arsenic-contaminated soil. Combined proteomic and metabolomic analysis of earthworms, exposed under laboratory conditions to natural soil spiked with arsenic, documented osmotic stress and immune dysregulation, distinguishing the abundances of asparagine and dimethylglycine as potential toxicity biomarkers - endpoints far more sensitive than conventional earthworm ecotoxicology assays. This molecular resolution can separate the distinct toxicity signatures of different contaminants. Earthworm transcriptomic and metabolomic data, combined with gut microbiome profiling, differentiated polymer-specific toxicity responses in Eisenia fetida exposed to different types of microplastics in agricultural soil under biogas slurry irrigation (Zhao, Y. et al., 2024). The gut community shifted in a polymer-dependent manner, mirroring the earthworms’ different stress responses to biodegradable and conventional MPs. Biodegradable PLA induced neurotransmission disorder and high energy expenditure in earthworms, while conventional PVC and LDPE activated immunity pathways and reduced nutrient absorption.
Analyzing multi-omic datasets is facilitated by specialized bioinformatics tools that resolve correlations across omic data types. Boughattas et al. (2024) integrated 16S gut microbiome data with metabolomics through the DIABLO framework (Data Integration Analysis for Biomarker discovery using a Latent component method for Omics studies; Singh et al., 2019) in the mixOmics package (Rohart et al., 2017), linking specific metabolites to microbial taxa within Eisenia andrei exposed to 2,4-dichlorophenoxyacetic acid, microplastics, and their combination. The co-contaminant scenario produced the most pronounced disruptions, affecting metabolites tied to oxidative stress, inflammatory response, amino acid synthesis, energy, and nucleic acid metabolism. Integrating molecular layers through multi-omics clarifies how contaminants cause toxicity, supporting more accurate risk interpretation.
Multi-omic studies of soil, plant, or fauna microbiomes provide rich biomarker libraries for TRIAD, leveraging bioinformatics software to determine subtle relationships across datasets. Using DIABLO within mixOmics, Romdhane et al. (2019) correlated shifts in the soil bacterial community with soil metabolome changes at increasing doses of two β-triketone herbicides. At ten times the agronomic dose, leptospermone disturbed bacterial diversity more strongly than sulcotrione, and DIABLO resolved a distinct OTU-metabolite signature for each herbicide. These herbicide-specific biomarkers, sought by the researchers as sensitive risk indicators, capture molecular microbial effects overlooked by regulatory assessment. Testing the response of long-term PAH-contaminated agricultural to fresh PAH input, Li, X. et al. (2019) combined data from soil enzyme assays, metabolomics, and PICRUSt (Phylogenetic Investigation of Communities by Reconstruction of Unobserved States; Langille et al., 2013), which predicts functional gene content using microbial taxonomy data. The fresh PAH dose inhibited dehydrogenase and urease activity, while correlation analysis linked 43 altered metabolite abundances to microbial community shifts, pointing to inhibited fatty acid, carbohydrate, and amino acid pathways. These genetic and metabolic responses serve as biomarkers of PAH contamination, relevant to EU soil remediation goals (
European Commission, 2021). Combining 16S sequencing with untargeted metabolomics, Wu, C. et al. (2022) applied LEfSe (linear discriminant analysis effect size; Segata et al., 2011), which pinpoints the taxa most distinctive to each condition, alongside PICRUSt to plastic mulch-contaminated soil. The analysis documented plastic-enriched taxa linked to potential microplastic degradation pathways. The genus
Clostridium_sensu_stricto_1
correlated with the caprolactam degradation pathway, a signigicant route for microplastic breakdown, illustrating the value of these methods for both ERA and the discovery of novel remediators.
Microplastic contamination has emerged as a central application of integrated multi-omics for soil ERA. After a 120-day exposure of farmland, woodland, and wetland soils to plastic residues, specific shifts per soil type were observed in microbial community composition, C/N/P-cycling taxa, and metabolite profiles. Biodegradable PLA disturbed soil enzyme activity and nutrient content more than conventional polyethylene, with woodland soil showing pH changes that farmland largely resisted. This high-resolution data distinguished the vulnerabilities of different soils to contaminant effects, a critical consideration for site-specific TRIAD assessments (Dong et al., 2024).
Combining metagenomics, PICRUSt prediction, and root-exudate metabolomics under PBAT (biodegradable) and PE (nonbiodegradable) microplastic contamination, Han et al. (2024) documented dose-dependent suppression of microbial C/N/P/S-cycling and pakchoi growth under PBAT, while PE caused no significant phytotoxicity. Four PBAT-enriched degrader genera were negatively related to plant growth, pointing to toxic degradation intermediates. Causal modelling explained 87% of plant-growth variation, attributing harm to both direct MP toxicity and indirect effects through altered soil properties and microbial communities (Han et al., 2024). A companion study found PBAT degradation rate itself varies with soil type, driven by differences in the soil microbial community (Han et al., 2021), implying the harm from biodegradable plastics depends on local soil conditions. In peanut rhizosphere soil, integrated metagenomics and metabolomics revealed that polypropylene and rubber crumb microplastics disrupted N-cycling through distinct mechanisms. Polypropylene promoted denitrification while inhibiting nitrification, whereas rubber crumb suppressed all three N-transformation processes (Liu, Y. et al., 2023). The analysis linked specific altered metabolites to these N-cycling gene shifts, while root-cell membrane damage and oxidative stress, both more severe under rubber crumb, further inhibited nitrogen uptake. Notably, between biodegradable and conventional microplastic comparison, biodegradable MPs consistently posed greater ecological threat than conventional, a result requiring further high-resolution research with direct implications for regulatory risk assessment.
Multi-omics also offers transformative potential for TRIAD-aligned remediation strategies. Bioremediation of PAH-contaminated soils can inadvertently increase genotoxicity, the capacity to damage DNA, through accumulation of oxy-PAH transformation products more genotoxic than the parent compounds. Using integrated metagenomics and metabolomics, Jiménez-Volkerink et al. (2024) characterized a microbial consortium capable of degrading one such compound, benz(a)anthracene-7,12-dione (BaAQ), from creosote-contaminated soil. Genome-resolved analysis confirmed that the complete degradation pathway relied on a division of catabolic labor between consortium members, while community assembly and survival relied on their nutritional interdependencies. Important degradation genes had spread through the community via horizontal gene transfer. This molecular-level mapping of degradation roles and dependencies provides the mechanistic understanding needed to design next-generation bioremediation strategies, a core objective of higher-tier TRIAD assessment.
3. Conclusion
Conventional soil risk assessment leaves gaps in sensitivity, attribution, and mechanism that molecular omic methods are positioned to fill within a modernized TRIAD framework. These methods detect contaminant effects at sub-lethal exposures that conventional toxicity endpoints miss, supplying the sensitive ecological measurements that current assessment needs. These methods also resolve the origin of an effect, distinguishing contamination from natural abiotic stress, isolating the contribution of a single contaminant within a mixture, and detecting antagonistic or synergistic interactions between contaminants or environmental factors. Such attribution is central to accurate risk interpretation, since a Line of Evidence holds value only when an observed effect can be traced to its specific cause. Additionally, they offer indicators capable of tracking dynamic change, providing fine-grained temporal data to differentiate short- and long-term effects, addressing a further gap in conventional TRIAD tests.
A recurring theme is that molecular data require other evidence to yield a risk interpretation. Researchers come to toxicological conclusions by integrating omic measurements with soil physicochemical properties, contaminant concentrations, physiological responses, and conventional soil microbial parameters and ecotoxicological endpoints. These data fall under TRIAD's chemical, ecological, and ecotoxicological Lines of Evidence. This dependence on integration is precisely why -omics fits into the multi-LoE TRIAD structure. Cecchetto et al. (2023) demonstrate that omic endpoints can be resolved into quantifiable, scalable measurements, whereby they can enter the integrated risk (IR) calculation directly, contributing to each Line of Evidence. Their sensitivity and discrimination let researchers see response patterns that improve how evidence is weighted and combined, refining and modernizing the TRIAD IR calculation.
Omic methods expand the reference libraries of contaminant-specific biomarkers needed to standardize molecular endpoints and to train predictive machine learning models. They trace toxicity through trophic levels, uncovering both direct and indirect effects of contamination, and connect genetic potential or expression to realized metabolic functions. Untargeted approaches can pinpoint degrading or resistant species, genes, and consortia that inform bioremediation design, a stated objective of higher-tier assessment and EU soil policy. However, robust predictive models become transferable across different environmental conditions only after training on large, representative datasets and undergoing external validation. Obtaining such datasets and performing this validation, both time and labor-intensive, remains a challenge of model development, which must be addressed by the scientific ERA community.
Realizing the potential of omic bioindicators for TRIAD requires placing each method where it contributes most.
Figure 5 presents the modernized tiered TRIAD framework, positioning omic methods within the ecotoxicological and ecological Lines of Evidence and scaling them by tier. Molecular methods enter at tier 2, with established and accessible approaches such as metabarcoding and functional-gene microarrays exploring community level shifts, and organism-level metabolomics probing sublethal ecotoxicological effects. Tier 3 deepens the ecotoxicological investigation with transcriptomic, proteomic, and plant- and fauna-associated microbiome profiling. Within the ecological LoE, community-level untargeted sequencing, identifies shifts in taxonomic and functional structure, and can discover novel resistant or degrading species or communities. Metabolomic and metaproteomic standalone assays offer assessment endpoints which validate actual functional response.
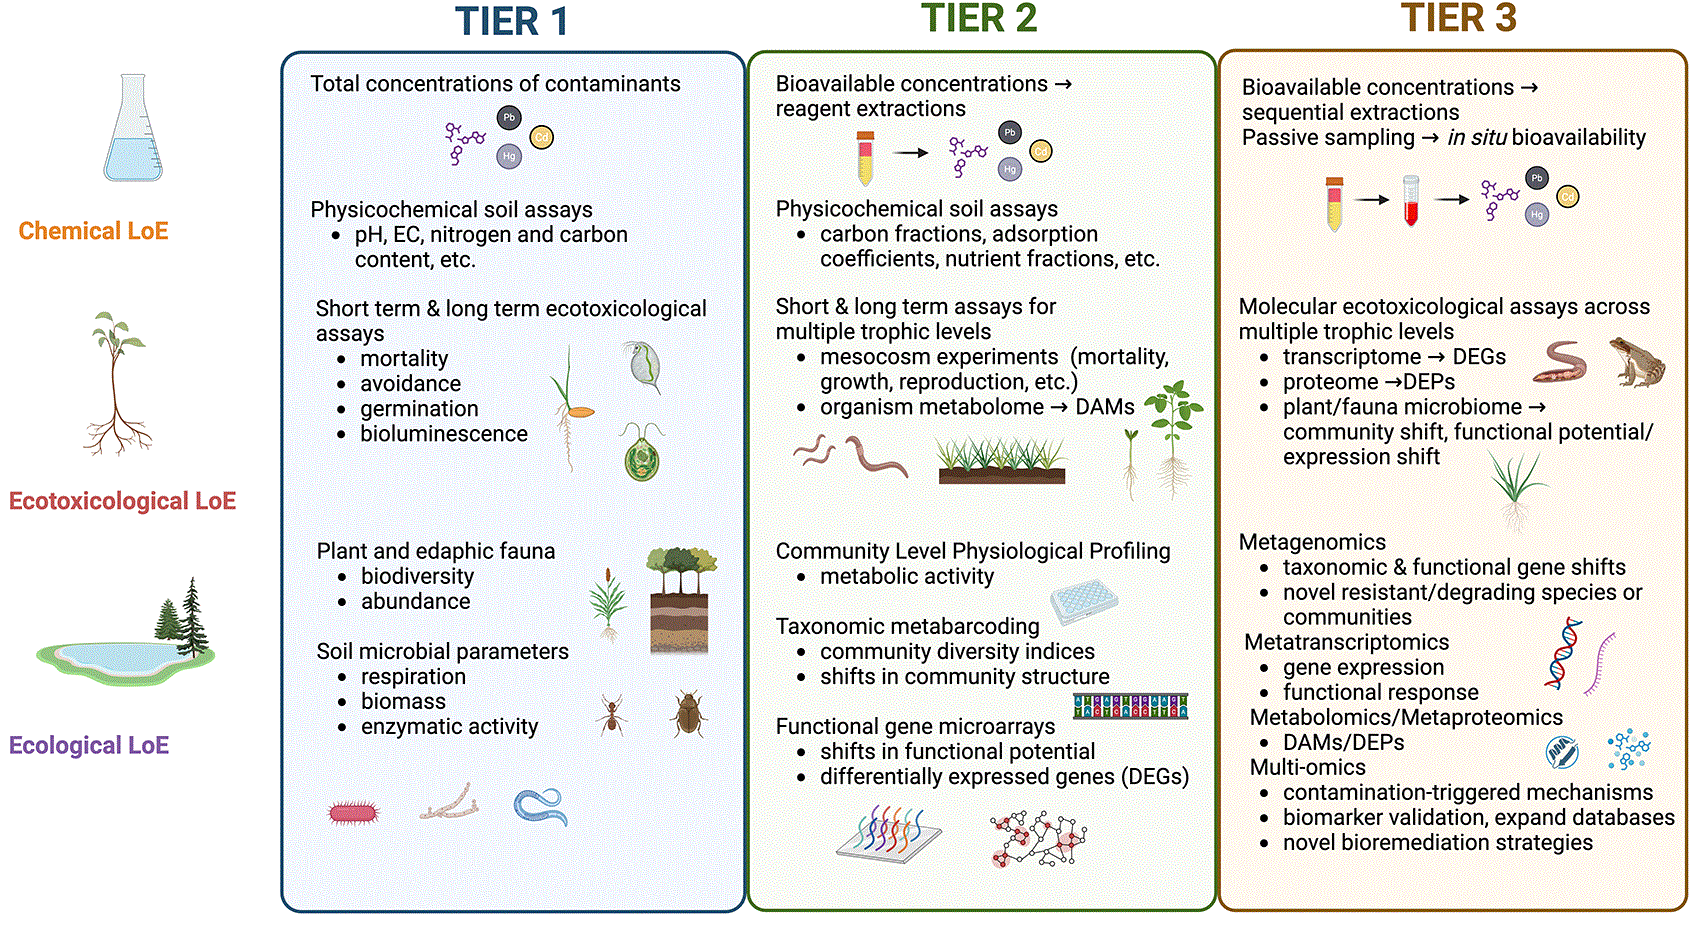
Figure 5. Molecular omic methods positioned within the modernized tiered TRIAD framework.
Representative tests and indicators are organized by the three TRIAD tiers (columns) and the chemical, ecotoxicological, and ecological Lines of Evidence (rows). Tier 1 takes on physicochemical and microbial soil quality indicators. Molecular methods begin at tier 2 with organism metabolomics for ecotoxicology (cheaper than proteomics); taxonomic metabarcoding, and functional-gene microarrays for the ecological LoE. Tier 3 adds the higher-resolution approaches: transcriptomics, proteomics, metagenomics, metatranscriptomics, metabolomics and metaproteomics, and integrated multi-omics for contamination-triggered mechanisms, biomarker validation, and bioremediation. Created with BioRender.
Integrated multi-omics are sensitive, multifunctional tools delineating contamination-triggered mechanisms, validating biomarkers, and guiding bioremediation design. This arrangement brings molecular resolution forward into routine mid-tier assessment while reserving the most resource-intensive analyses for tier 3, preserving the practicality of the TRIAD framework. Standardized protocols, reference databases, and structured and transparent scoring systems remain prerequisites for maintaining the TRIAD's contemporary relevance (Part I, Djomehri et al. 2026). The studies synthesized here establish that molecular endpoints supply the sensitivity, attribution, and mechanistic depth that soil risk assessment has lacked, reinforcing the ecological and ecotoxicological Lines of Evidence and, with them, the integrity of the integrated TRIAD risk estimate.
Future work should test these proposed integrations within published TRIAD datasets and operational contaminated-site assessments, determining how molecular endpoints affect line-of-evidence weighting, integrated risk estimates, and remediation decisions. Applied to real cases, this evaluation would supply the proof of concept the modernized framework now needs, refining analytical methods and generating the datasets that anchor and verify predictive models. It is through this grounding in practice that the modernizations proposed in Parts I and II, and the integration of omic evidence in particular, will move from proposal to established assessment tool.