1 Introduction
Particulate organic carbon (POC) is operationally defined as the non-carbonate and combustible carbon matter in particulate form that is retained by a filter of a typical nominal pore size of 0.7 µm (Kharbush et al., 2020, IOCCG POC Protocols). In the open-ocean, pelagic POC mainly includes suspended detrital matter (i.e., remains of dead organisms) and living organisms (i.e. phytoplankton, heterotrophic bacteria and zooplankton) (Gardner et al., 2006; Turnewitsch et al., 2007).
POC constitutes the third most abundant carbon pool (~2 Pg C, Brewin et al., 2021; Stramska & Cieszynska, 2015) in the ocean, the others being the dissolved inorganic (~38,000 Pg C, Hedges, 1992) and dissolved organic (~662 Pg C, Hansell & Carlson, 2013) and the particulate inorganic carbon pools (~0.03 Pg C, Hopkins et al., 2019). POC has one of the highest turnover rates among the pools of carbon in the ocean (Brewin et al., 2021; Sarmiento & Gruber, 2006) and the efficiency with which it is transferred from the upper to the deep ocean and sediments contributes to controlling atmospheric CO2 concentrations (Kwon et al., 2009; Parekh et al., 2006).
In the last decades, several studies have collected relatively extensive measurements of POC, leading to progress in understanding the nature of POC and the processes controlling its spatio-temporal distribution (Aumont et al., 2017; Bishop & Wood, 2008; Bishop & Wood, 2009; Cetinić et al., 2012; Gardner et al., 2003; Gardner et al., 2006; Henson et al., 2012; Kharbush et al., 2020; Kiko et al., 2017; Lam et al., 2011; Moran et al., 1999; Rasse et al., 2017; Turnewitsch et al., 2007; Wangersky, 1976). However, even though much effort has been invested in developing methods for determining POC (Gardner et al., 2000; Gardner et al., 2003; Gardner et al., 2006; Liu et al., 2005; Liu et al., 2009; Moran et al., 1999; Wangersky, 1974; Wangersky, 1976), we still have a relatively rudimentary understanding of the uncertainties associated with the different steps of the analysis and, therefore, with the resulting POC estimates. This is a problem because identifying and quantifying such uncertainties will ultimately allow the community to further understand how and why POC varies in the ocean. In addition, by better characterising POC uncertainties and their major sources, we might improve the method for determining POC.
The objectives of this study were 1) to present a method to experimentally quantify the uncertainties of POC measurements based on duplicate filters in both pelagic and deep ocean layers; 2) to model POC uncertainty based on assumed sources of uncertainty affecting the determination of POC; and 3) to compare modelled and experimental uncertainties. Our results show that modelled uncertainties accounted for only a small fraction of the experimental POC uncertainties, suggesting that sources of uncertainty different from those considered in our analysis controlled the uncertainty of our POC determinations. Identifying, characterising and minimising these additional uncertainties will lead to improved measurements of POC and a better understanding of its variability and the role it plays in the ocean carbon cycle.
2 Data and methods
At the time when the data presented in this study were collected, the best known and accepted protocol for determining POC was the one established in the mid 90’s for The Joint Global Ocean Flux Study (JGOFS) (Knap et al., 1996). Studies that appeared after the JGOFS protocol identified additional potentially important sources of uncertainty in the POC determination, as for example the adsorption of DOC onto the POC filters or the volume of water needed to minimise uncertainties when POC values are low (Cetinić et al., 2012; Gardner et al., 2003; Gardner et al., 2006; Moran et al., 1999; Stramski et al., 2008). In an attempt to quantify and minimize the uncertainties of the POC concentrations determined in the present study, we modified the sampling, processing and analysis described in the JGOFS protocol. Modifications, described in more detail in Section 2.1, Section 2.2 and Section 2.3, included the sampling of different volumes of water in accordance to expected in-situ concentrations, and using different types of blanks to quantify the dissolved organic carbon and any contamination due to the acidification step. Yet, minimising all uncertainties proved difficult. More recently, a NASA-led team has been developing a revised and considerably more detailed POC protocol (currently in draft version) to support the validation of ongoing and upcoming ocean-colour satellite missions (see IOCCG POC Protocols).
2.1 Sampling
Water samples were collected at 68 stations during the 24th Atlantic Meridional Transect (AMT-24) aboard the RRS James Clark Ross from September 25th to November 1st, 2014 (see Figure 1). Two casts were completed every day (weather permitting): one pre-dawn and the other around solar noon. Different ecological provinces (Longhurst, 2007) were sampled: the North Atlantic Drift Province (NADR), the North Atlantic Subtropical Gyral Province (NAST), the North Atlantic Tropical Gyral Province (NATL), the Western Tropical Atlantic Province (WTRA), the South Atlantic Gyral Province (SATL), and the South Subtropical Convergence Province (SSTC).
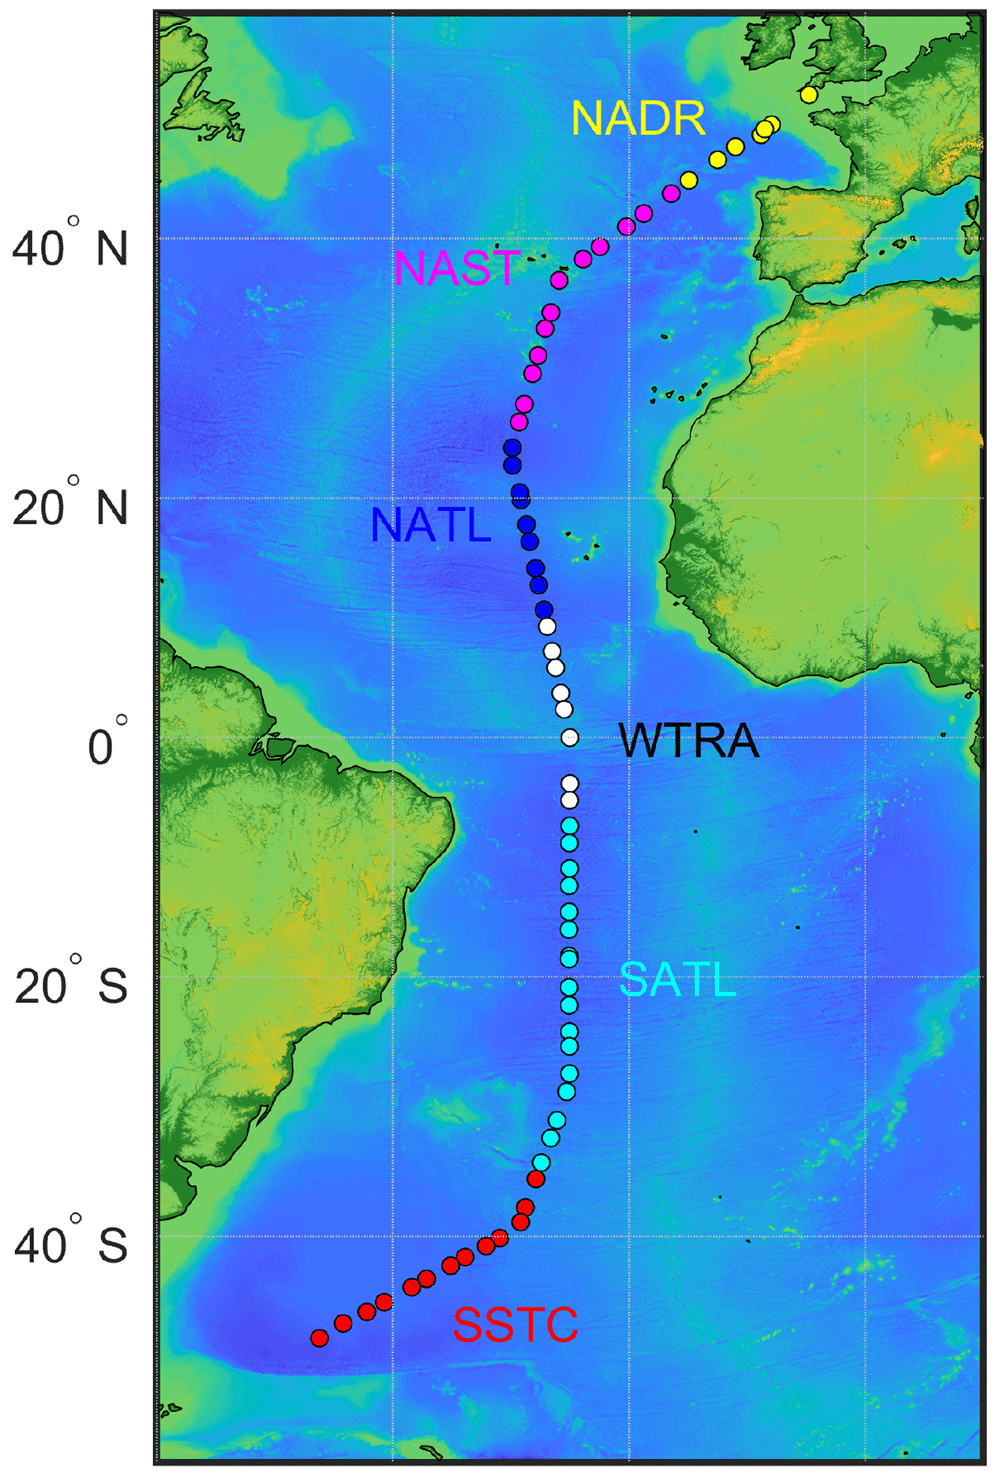
Figure 1. Track of AMT-24 cruise with the locations of 67 stations where samples for particulate organic carbon concentration were collected.
Colour coding of the stations represents biogeographical provinces that were sampled: the North Atlantic Drift Province (NADR), the North Atlantic Subtropical Gyral Province (NAST), the North Atlantic Tropical Gyral Province (NATL), the Western Tropical Atlantic Province (WTRA), the South Atlantic Gyral Province (SATL), and the South Subtropical Convergence Province (SSTC).
At each station, water samples were collected from six depths in the upper 500 m (with the exceptions of stations 13 and 43 where 3 and 5 samples were collected, respectively, and of stations 23 and 24 where all samples were lost) using 20-l Niskin bottles (Ocean Test Equipment Inc., Standard ES, Model 115) that were installed on a bespoke stainless-steel frame. The sampling procedure did not account for dregs, the rare large particles that might have been under sampled by Niskin bottles (Gardner et al., 2006). Water samples were transferred from the Niskin bottles into six 15-l HDPE carboys, which were pre-washed with 10% HCl and then with sample water prior to use. The carboys were taken to the on-board laboratory, where water samples were transferred into six ~2-l narrow-mouth amber bottles (Thermo Scientific Nalgene) and filtered through pre-combusted (450°C for five hours) 25 mm Whatman glass fiber filters (GF/F, nominal pore size 0.7 µm) at low vacuum (~125 mm Hg) by inverting the bottles into a standard funnel setup. Each of the bottles was also pre-washed with sample water prior to use. To keep the water samples homogenised, carboys were gently shaken before pouring into the bottles. The volume of sample water filtered for each POC measurement varied between ~1 and ~8 liters, and it was adjusted according to the expected concentration of POC in the respective environment. Thus, each 2-l bottle was typically re-filled (up to four times) during filtration to achieve the total volume for any given sample. No measures were taken to prevent atmospheric contamination during filtration.
We implemented the "double-filter" technique advocated by Kinney et al. (1971); Banoub and Williams (1972); Loder and Hood (1972); Feely (1974); Smith et al. (1996); Moran et al. (1999). These authors suggested using two stacked filters for each estimate of POC concentration. The upper filter was used to collect particles and adsorbed dissolved organic carbon (uncorrected POC sample, henceforth referred to as "uPOC") while the lower filter quantified the dissolved organic carbon adsorbed onto GF/F filters (adsorbed DOC blank, henceforth referred to as "aDOC") (see Table 1 for a list of variable names and their subscripts). After each filtration, uPOC and aDOC filters were removed from the filtration rig, wrapped into separate pre-combusted (450°C for five hours) aluminium foil envelopes, flash-frozen in liquid nitrogen, and then stored in a freezer at −80°C for post-cruise analysis.
Table 1. List of abbreviations and symbols.
| Subscript | Variable Name | Units |
|---|
| POC | Particulate Organic Carbon | N/A |
| uPOC | Uncorrected Particulate Organic Carbon | N/A |
| aDOC | Adsorbed Dissolved Organic Carbon | N/A |
| MuPOC | Mass of organic carbon on uPOC | μg |
| MaDOC | Mass of organic carbon on aDOC | μg |
| Mcap | Mass of organic carbon on tin capsules | μg |
| Mac | Mass of organic carbon on acidified filters | μg |
| Mnac | Mass of organic carbon on non-acidified filters | μg |
| Blank-corrected mass of organic carbon on uPOC | μg |
| Blank-corrected mass of organic carbon on aDOC | μg |
| M | Mass of particulate organic carbon | μg |
| C | Concentration of particulate organic carbon | mg/m3 |
| D1 | Carbon concentration of the first duplicate | mg/m3 |
| D2 | Carbon concentration of the second duplicate | mg/m3 |
| Average of POC concentration in the two duplicates | mg/m3 |
| Δ | Scaled arithmetic difference between duplicate pairs | mg/m3 |
| Δr | Scaled relative difference between duplicate pairs | dimensionless |
| σr | Relative experimental uncertainty of POC estimates | mg/m3 |
| σC | Absolute experimental uncertainty of POC estimates | mg/m3 |
| σV | Uncertainty in volume | L |
| σM | Uncertainty of carbon mass predicted by the calibration equation | μg |
| σC(V) | Modelled uncertainty in C due to uncertainty in volume | mg/m3 |
| σC(M) | Modelled uncertainty in C due to the uncertainty in calibration | mg/m3 |
| σC(η) | Modelled uncertainty in C due to the uncertainty in sample handling | mg/m3 |
| LC | Critical value | μg |
| LD | Detection limit | μg |
At each cast, a duplicate sample from a randomly chosen depth was collected to assess the uncertainty of our method, starting at station 8 with the exception of stations 23, 24, 39, 45, and 46 were duplicates samples were not collected. At station 43 duplicates at two depths were collected. In total, we collected 392 uPOC samples with their corresponding aDOC blanks and 57 duplicate uPOC samples with their related aDOC blanks. Sample water for each pair of duplicate measurements was taken from the same Niskin bottle. A larger number of replicates per sample would have provided more robust estimates of the POC uncertainty. Yet, in our case duplicates were chosen as a compromise between statistical robustness, the water available from the rosette for our analyses, and the time required to collect and analyse the samples. Thus, the statistics of the population of duplicates were used to determine an overall-cruise estimate of the relative POC experimental uncertainty (see Section 2.4.1).
2.2 Laboratory sample handling
All the filters were processed in 16 separate batches or CHN runs. Each run consisted of (1) uPOC filters and their corresponding aDOC blanks from multiple casts, (2) duplicate uPOC and aDOC filters from these casts used to estimate total experimental uncertainties, (3) empty tin capsules, acidified and non-acidified filter blanks (detailed below) used to estimate uncertainties related to the sample handling in the lab and systematic biases, and (4) standards used to calibrate the CHN analyser (see Section 2.3 for details).
Acidification at room temperature for a period of 12 to 16 hours was used to remove the inorganic carbon accumulated on the uPOC and aDOC filters. To do this, a crucible containing a small amount of 37% HCl (Sigma Aldrich, High Purity, 08256-500ml F) was located in the middle of a glass desiccator. The uPOC filters and aDOC blanks from each batch were removed from their aluminium envelopes, placed into individual acid-washed glass vials, and positioned around the crucible in the desiccators. In contrast to adding an aliquot of a dilute acid solution directly onto the filters, the technique of exposing them to acid fumes is expected to homogenize the effect of the acid on all the filters within a desiccator and to avoid losses of organic particles (Martin et al., 1993). To minimise differences in contamination between corresponding uPOC and aDOC samples, paired uPOC and aDOC filters were acidified in the same desiccator. Duplicate filters were acidified in different desiccators.
To account for any potential contamination during acid fuming, we introduced an additional type of pre-combusted (450°C for five hours) 25 mm Whatman GF/F filter blank. Three of these blank filters were processed as the sample filters and subjected to acid fuming in the desiccator, while three other blank filters were kept clean and dry outside of the desiccators.
After the acidification phase, all the filters, including the acidified uPOC filters, aDOC blanks, and acidified and non-acidified filter blanks were dried in an oven at 60°C for several hours. Subsequently, the acidified and non-acidified filters and blanks were wrapped into individual tin capsules (Pressed, Standard Clean, 10 × 10 by OEA Labs) and analysed for carbon. An increment of the C mass on the acidified filter blanks in comparison with the non-acidified ones would indicate contamination during sample acidification and drying.
2.3 Determination of POC
The mass of carbon contained on filters was determined by high-temperature combustion (Gordon & Sutcliffe, 1974; Menzel & Vaccaro, 1964; Sharp, 1974; Wheeler et al., 1997) using a CHN analyser (FlashEA 1112 Elemental Analyser, with helium CP grade N5.0 as carrier gas). Filters were processed in accordance with the manufacturer’s manual (ThermoQuest, 1999). In every CHN run, a new combustion tube was used. The extracted CO2 was measured by a thermal conductivity detector.
As samples are analysed, the reaction tube of the FlashEA CHN analyser gradually fills up with the combusted tin capsules and filters. Thus, as the CHN run proceeds and the reaction tube fills up, a variation in the instrument combustion efficiency can occur with time. To stabilise the combustion efficiency of the instrument throughout each run, we performed constant adjustments to the “sample delay” parameter, which represents the time that it takes the CHN analyser to combust each sample, to deliver CO2 released from the sample to the detector, and to run the analysis.
The CHN analyser was calibrated during each run using two sets of pre-weighted (Sartorius MC5 high-accuracy microbalance, calibrated yearly) acetanilide standards (C = 71.09%, N = 10.36%, OEA Labs, R66005) contained in tin capsules. The first set of 11 standards covered the entire range of expected masses of carbon on our uPOC and aDOC filters (5 - 300 µg) and it was analysed immediately prior to processing the sample filters. We will refer to this set of standards as the calibration standards. The second set of standards was processed alongside the filters (one standard after every six filters) to validate the initial calibration throughout filter processing. We will refer to the second set of standards as the stability standards. We note that, filters are expected to affect the combustion process and thus the sensitivity of the analyser (Planchon, personal communication, 2022). As a consequence, to reduce uncertainties and improve sensitivity, future studies should consider investigating how the uncertainty of POC varies when standards are first collected on clean filters and then treated as samples, rather than simply added to tin capsules.
In the absence of any instrumental drift, we expected the calibration coefficients derived from both sets of standards from the same CHN run to be statistically indistinguishable. However, during two out of 16 runs the CHN analyser was unstable for unknown reasons and the calibration coefficients derived from the two sets of standards differed significantly. Thus, we decided to use both types of standards, calibration and stability, to develop the relationship between the response of the CHN analyser and the mass of carbon on processed filters. To better characterise the instrument’s behaviour during the analysis, future work should also use standards with different weights at the end of the CHN run.
The mass of carbon M on the ith filter, processed during the kth CHN run, was estimated by a linear regression model using a robust fitting method that minimises the impact of outliers on the derived regression coefficients ("iteratively reweighted least-squares" implemented in the Matlab function fitlm as option RobustOpts):
where x represents the output signal from the CHN analyser, m and b represent the slope and the intercept of the regression line, respectively. The intercept was removed from the model when it was not statistically significant (p-value > 0.05).
For each CHN run, we estimated the mass of organic carbon contained on uPOC (MuPOC), aDOC (MaDOC) filters, tin capsules (Mcap), and acidified (Mac) and non-acidified (Mnac) GF/F filters using Equation 1. These MuPOC and MaDOC values, however, do not represent the true load of particulate organic carbon contained in the corresponding water samples as these values may be affected by biases, i.e., contamination during the acidification step, residual organic carbon on the tin capsules, and on the combusted GF/F filters). Therefore, the blank-corrected mass of organic carbon from the ith uPOC and aDOC filters ( and , respectively), which were acidified together in the jth desiccator and processed in the kth CHN run ( and , respectively), must be written as (notation as in Turnewitsch et al., 2007):
where is the average carbon mass of the three filter blanks acidified in the same desiccator jth as the ith filter and and are the average carbon mass of the three non-acidified filter blanks and the average carbon mass of three tin capsules, respectively.
For each pair of uPOC and aDOC filters, the mass of POC, M, was determined as
We assumed that uPOC and aDOC filters had adsorbed the same amount of DOC, and their contamination due to sample handling during the CHN analysis was equal to the average mass of the three acidified filter blanks. Hence, the subtraction in Equation 4 removed various systematic biases from the final estimates of the mass of POC. Note that because we processed corresponding pairs of uPOC and aDOC filters in the same desiccator and during the same CHN run, Equation 4 is equivalent to:
To determine the POC concentration, C, for each water sample, we divided M by the volume of water, V, filtered for each sample:
2.4 Uncertainty analysis
The standard law of propagation of uncertainty (JCGM, 2008) was used throughout our uncertainty calculations and we recall it here for the reader:
where is the total (or "combined") variance of the estimate y (POC, in our case), which is determined from the input quantities x1, x2, ..., xN through the functional relationship y = f (x1, x2, ..., xN). The uncertainty of each of the input variables is denoted as and their inter-dependencies are represented by the correlation coefficients r(xi,xj). The total uncertainty is the positive square root of .
2.4.1 Experimental uncertainty. We first estimated the overall experimental uncertainties associated with our POC concentrations by analysing the duplicate samples. These experimental uncertainties are expected to represent the uncertainties arising from all (or at least most of) the steps required to estimate POC concentrations (i.e., from sample collection in the field to sample analysis in the laboratory). Statistics from the duplicates differences were then used to estimate “whole-dataset” uncertainties, rather than per-sample uncertainties.
Absolute experimental uncertainties were estimated from the scaled arithmetic differences between the POC concentrations of the duplicates, D1 and D2 (Hyslop & White, 2009):
Since both D1 and D2 are uncertain and their uncertainties add in quadrature, to estimate the uncertainty in only one measurement, the difference of the duplicates was divided by (Hyslop & White, 2009).
However, the duplicate differences Δ were positively related to POC concentration and this dependency varied between the productive (correlation coefficient r = 0.40, p = 0.0009) and mesopelagic (r = 0.54, p = 0.0005) zones (Figure 2a). To remove this dependency on concentration, we expressed the differences in duplicate measurements as a relative difference (Δr = Δ/), where is the average of POC concentration in the two duplicates. Figure 2b confirms that, once normalised, the relative duplicate differences in the productive and mesopelagic zones did not depend on POC anymore.
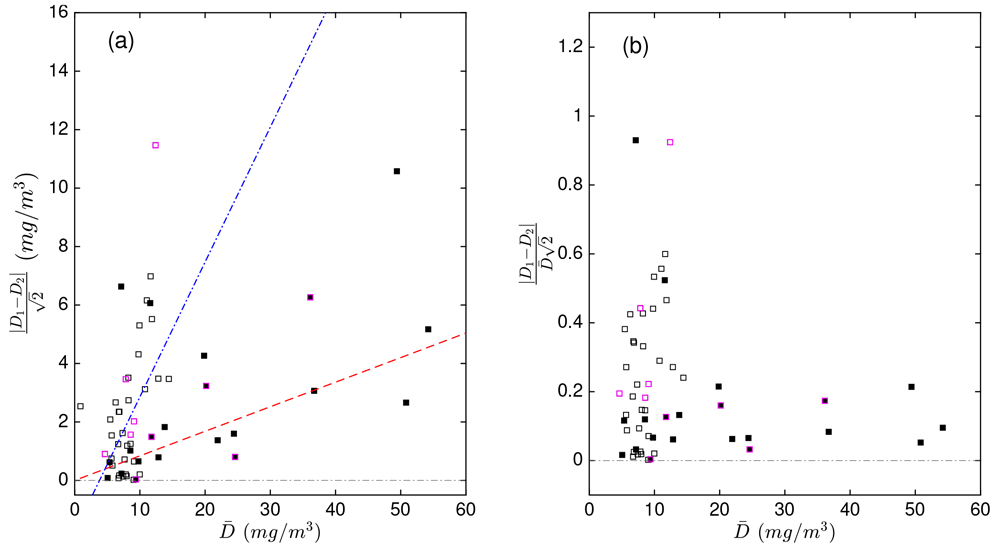
Figure 2.
Scaled differences in duplicate particulate organic carbon (POC) measurements (a) as a function of the mean value of each pair of duplicates (). Scaled relative differences in duplicate POC measurements relative to (b). The absolute values were used to more easily demonstrate the dependency of the duplicate differences on POC. Black and white points represent duplicate measurements from the productive and mesopelagic zones, respectively. Points with magenta borders represent samples processed during CHN runs with highly uncertain calibrations. The red dashed and blue dash-dotted lines are the linear fits to the data in the productive and mesopelagic zones, respectively.
To estimate “typical’ relative uncertainties in POC (σr) that can then be applied to the entire dataset, we needed to minimise the influence of outliers in the distribution of Δr (Hyslop & White, 2009). Therefore, σr was estimated as the robust standard deviation of the relative duplicates in each zone:
where P84 and P16 are the 84th and 16th percentiles of Δr (Hyslop & White, 2009).
Finally, the absolute experimental uncertainty of POC concentration was estimated for each sample as:
2.4.2 Modelled uncertainty and uncertainty budget. In this section, we used a second method to model an independent estimate of the total uncertainty in POC concentrations. Specifically, the standard law of propagation of uncertainty was used to propagate the uncertainties associated with different steps of the POC determination. This independent model estimate of the POC uncertainty was then compared to the total experimental uncertainty derived from the duplicate measurements to assess the extent to which this latter theoretical calculation could reproduce the experimental uncertainties: the closer these estimates are, the more confident we can be in how we understand the measurement process and its uncertainties.
Equation 7 also allows one to estimate an uncertainty budget, that can be used to partition the total uncertainty into different contributions. Here, the relative contribution ux of each modelled uncertainty source σC(x) was computed as the ratio of the uncertainty of each specific modelled uncertainty source divided by the total experimental uncertainty:
The different relative uncertainty contributions can be ranked to identify the most uncertain steps in the methodology and prioritise improvements in the method. Finally, the sum of the modelled relative variances can be compared to the total experimental variance to quantify the fraction of experimental variance that we were not able to model (i.e., explain).
Uncertainty in POC concentration due to uncertainties in the sample volume
Errors in measuring the volume of sample seawater translate into uncertainties in POC concentration. Since each POC sample required from one to five bottles of sample seawater, each with a volume Vn, the combined uncertainty of the total volume, V, of seawater used for a sample depended on the number of bottles, n, used and the uncertainty in volumetric measurements, σVn, of each bottle. σVn was set equal to half of a graduation mark of the measuring cylinder (uncertainty of 10 ml). The volume of each bottle (~2.2 l) was measured with a measuring cylinder multiple times. Thus, the combined uncertainty of volume V can be expressed as
We estimated the uncertainty in POC concentration due to the uncertainty in volume, σC(V), by applying the propagation of uncertainty to Equation 6 and obtained:
Uncertainty in POC concentration due to uncertainties in the calibration equation
The uncertainty in the calibration equation (Equation 1) was expected to be one of the largest contributors to the uncertainty budget. To estimate this uncertainty, we first estimated the uncertainties σM of the aDOC and uPOC carbon masses predicted by our calibration equation. We did this by using 68% prediction intervals (PIs, the 68% was selected to conform with the common notion of one standard deviation). A PI is defined similarly to the better known confidence interval. However, the prediction interval is more appropriate for quantifying the uncertainty of the calibration equation because it estimates the expected uncertainty of an individual future observation by taking into account the uncertainties arising from all the regression parameters (Rawlings et al., 1998).
For each CHN run, we estimated σM as:
where σres represents the robust standard deviation of the residuals of carbon mass about the calibration equation, x is the instrument response (Equation 1), and sx are the mean and the standard deviation of x, nS is the number of standards used to fit the model, and t1−α/2, is the value from the t distribution with nS − 2 degrees of freedom and α equal to 0.32 corresponding to a 68% prediction interval (Altman, 2000).
The uncertainty of the calibration equation depended on both the uncertainty in the weights of the standards and the sensitivity of the instrument at the time when filters were processed. Therefore, σres and σM varied among CHN runs. Two CHN runs had significantly higher σres and therefore generated higher uncertainties in mass estimates, compared to the other runs. We believe that this was because of a less precise weighting of the standards, and not an instability or lower sensitivity of the CHN analyser. In support of this hypothesis, we found that, on average, the differences of the duplicate POC estimates derived from these specific calibration equations were not larger than those derived from calibration equations with lower uncertainties (Figure 2). This means that there was no bias in the less precise standards - they resulted in unbiased calibration coefficients, even though the random uncertainties of these coefficients were higher. Nevertheless, to avoid skewing our uncertainty estimates towards higher values, we did not include data derived from these two CHN runs when we estimated the POC experimental uncertainty (Equation 10).
We then estimated the uncertainty in POC concentration due to the uncertainty in the calibration equation, σC(M), by propagating the uncertainties σMuPOC and σMaDOC to the uncertainty of POC concentration by applying Equation 7 to Equation 4 and Equation 6:
where r (MuPOC, MaDOC) is the correlation coefficient between MuPOC and MaDOC.
Uncertainty due to laboratory contamination
During laboratory analyses aimed at quantifying carbon masses on filters, blank and sample filters can be contaminated resulting in biases and/or increased uncertainty of the estimates of POC concentration (King et al., 1998). This uncertainty can be introduced during filter-handling steps such as thawing, acidification, drying, and encapsulation. Considering that the uPOC samples, their corresponding aDOC blanks, and the filters blanks were acidified in the same desiccator and dried together in the oven, the amount of contamination that they might have received should have been approximately equal. Thus, any potential bias due to this contamination should have been minimised when was subtracted from in Equation 4. However, the carbon masses of the three acidified filter blanks for each desiccator varied slightly, indicating either that (i) the mass of organic carbon remaining on the pre-combusted filters and/or tin capsules could have varied or (ii) that the estimated contamination was uncertain or (iii) both.
The uncertainty in POC concentration due to uncertainty in contamination during laboratory analysis, σC(η), was estimated by applying the standard law of propagation of uncertainty to Equation 6:
where σηu and σηa represent the uncertainties of contamination due to handling of uPOC and aDOC filters, respectively. These uncertainties were assumed to be equal for uPOC and aDOC filters (i.e., ση) and were estimated as the standard error of the mean of the carbon masses of the three acidified filter blanks in each desiccator.
2.5 Detection limits
To determine the limit of detection of our technique we used the approach recommended by The International Union of Pure and Applied Chemistry (IUPAC) and The International Organization for Standardization (ISO) (Analytical Methods Committee AMCTB No. 92, 2020). We calculated the detection limit, (LD), by first computing the critical value, (LC, Equation 17), which establishes the presence of the analyte (carbon in our case), and is defined as the minimum significant estimated value of an analytical result, which is used as to discriminate against background noise (Currie, 1995):
where and s0 are the mean and the standard deviation of a blank material free from carbon, in our case, the tin capsules. t0.95;d f is the one-tailed 95% quantile for Student’s t with degrees of freedom d f, according to the number of values used to estimate and s0.
Given LC, we estimated LD, which is defined as the lowest carbon mass that our analytical method is reliably capable of detecting 95% of the times:
2.6 Reporting uncertainties
To estimate uncertainties associated with median values reported throughout the manuscript, we used the robust standard deviation (as described by Equation 9, after substituting Δr with the appropriate variable for which the uncertainty is being estimated).
3 Results and discussion
3.1 Distribution of POC
After excluding data from the unstable CHN runs (see section 2.3), the number of samples was reduced to 190 and 134 for the productive and mesopelagic zones, respectively (Table 2). The POC concentrations from the AMT-24 cruise were highly variable, ranging between 2 and 76 mg/m3 (Figure 3). The overall median (± 1 robust standard deviation) POC concentration from the productive zone was 19(±20) mg/m3, whereas that from the mesopelagic zone was 7(±3) mg/m3. The uncertainties associated with the median values represent the spatial variability along the transect. We also observed latitudinal patterns of POC driven by the seasonality and differences between the biogeographical regimes sampled during the cruise (Figure 1 and Figure 3). In the productive zone, the highest POC concentrations were found at temperate latitudes and around the equator, whereas the most oligotrophic provinces were characterised by lower POC concentrations (Table 2). Such distribution in the POC concentration matched the typical latitudinal patterns encounter for the upper layer of the Atlantic ocean (Poulton et al., 2006; Rasse et al., 2017; Wangersky, 1976). Particularly, high POC concentrations were found in the sub-surface (50 m) of the South Subtropical Convergence Zone. In this province, Poulton et al. (2006) found relatively high POC concentrations deeper than 200 m as a result of sinking particles from the euphotic zone of the SSTC. Overall, POC concentration below the productive zone was less variable throughout the transect, but the latitudinal pattern remained (see Table 2).
Table 2. POC concentrations for productive and mesopelagic zones across the sampled biogeographical provinces: the NorthAtlantic Drift Province (NADR), the North Atlantic Subtropical Gyral Province (NAST), the North Atlantic TropicalGyral Province (NATL), the Western Tropical Atlantic Province (WTRA), the South Atlantic Gyral Province (SATL), and the South Subtropical Convergence Province (SSTC).
"Std" are robust standard deviations and represent spatial variability. Median and Std values are expressed in mg/m3. Samples analysed during the unstable CHN runs were excluded.
| Productive zone | Mesopelagic zone |
|---|
| Province | Samples | Median | Std | Samples | Median | Std |
|---|
| NADR | 19 | 49 | 28 | 10 | 7 | 3 |
| NAST | 7 | 27 | 11 | 6 | 6 | 1 |
| NATL | 36 | 14 | 10 | 18 | 8 | 2 |
| WTRA | 32 | 19 | 10 | 16 | 9 | 4 |
| SATL | 57 | 14 | 6 | 44 | 7 | 2 |
| SSTC | 39 | 46 | 23 | 40 | 9 | 3 |
| ALL | 190 | 18 | 9 | 134 | 7 | 2 |
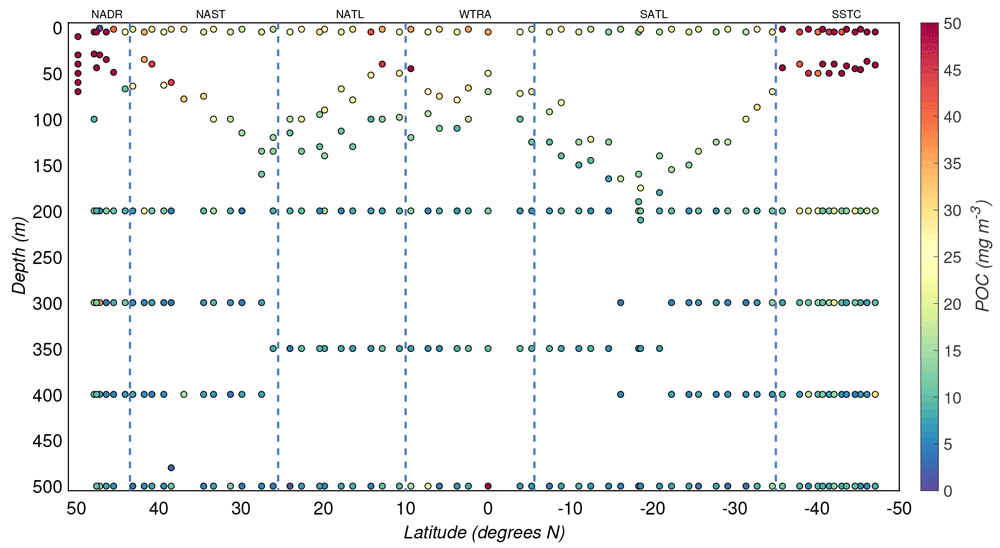
Figure 3. Depth-resolved distribution of particulate organic carbon (POC) concentration along the AMT-24 cruise.
Borders of the sampled biogeographical provinces are marked by blue vertical lines.
Even though our POC concentrations are in the range of those published in the literature, a direct comparison is complicated due to differences in methodologies, sampling times, and regions. The range of POC concentration estimated from bottle samples in the Atlantic ocean by numerous researchers spans from 5 to 350 mg/m3 (Balch et al., 2010; Banoub & Williams, 1972; Cetinić et al., 2012; Gardner et al., 1993; Gardner et al., 2003; Gardner et al., 2006; Graff et al., 2015; Marra et al., 1995; Menzel & Goering, 1966; Mishonov et al., 2003; Poulton et al., 2006; Stramska & Stramski, 2005; Stramski et al., 2008; Wangersky, 1974; Wangersky, 1976). A limited number of studies present POC concentrations measured in the Atlantic ocean between 100 and 500 m and spanning from ~0 to 35 mg/m3 (Carlson et al., 2000; Cetinić et al., 2012; Menzel, 1967; Poulton et al., 2006; Wangersky, 1974; Wangersky, 1976).
3.2 Detection limits
The median carbon mass from all the tin capsules used in the 16 CHN runs was 2(±1) µg. Thus, the estimated LC and LD were 3 and 5 µg C, respectively (Figure 4). The vast majority of mesopelagic aDOC filters collected and analysed during this study were above LD, with just four filters falling below LD.
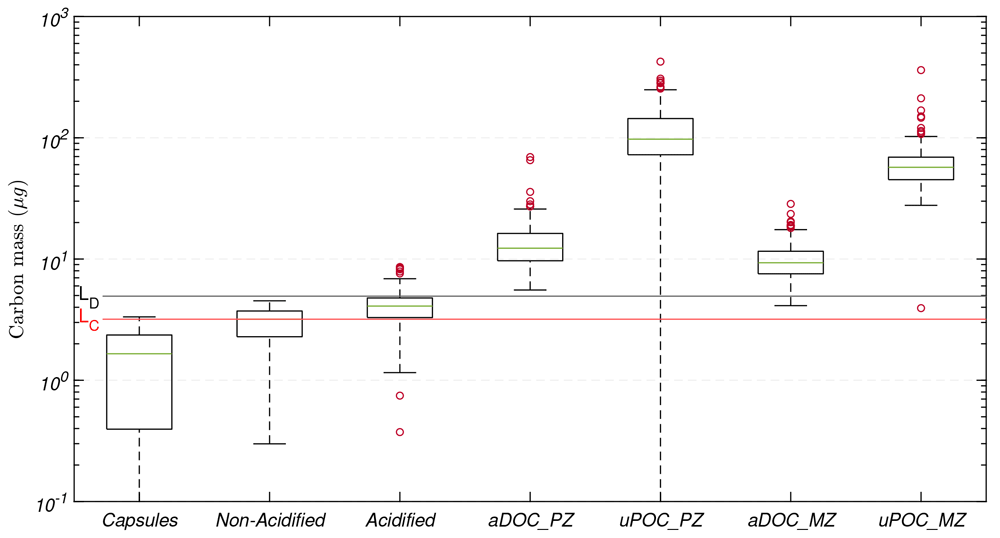
Figure 4. Distribution of carbon mass determined on tin capsules, non-acidified and acidified blank filters (GF/F) used for sample processing, and adsorbed dissolved organic carbon (aDOC) and uncorrected particulate organic carbon (uPOC) filters.
In each element of the box plot, the central rectangle spans from the first quartile (25th percentile) to the third quartile (75th percentile). The green line inside each rectangle shows the median value and whiskers below and above the box show the locations of the 5th and the 95th percentile, respectively. Red circles represent outliers. Red and black horizontal lines represent the critical value (LC) and detection limit (LD), respectively.
3.3 Correction for biases
We corrected additional sources of bias in the estimates of POC concentration by subtracting aDOC blank measurements from the corresponding uPOC measurements (see Section 2.3 and Equation 4). Since the carbon mass determined on an aDOC blank includes the mass of the adsorbed DOC, the carbon masses detected on empty tin capsules, clean GF/F filters and any contamination occurring during filter acidification and handling, it represents the cumulative bias for which the carbon mass on a corresponding uPOC filter needs to be corrected. Hence, to minimise biases introduced by any potential contamination and mass predicted by the calibration equation, we processed pairs of uPOC and aDOC filters together during sampling, acidification, handling, and processing stages, i.e., acidified in the same desiccator and analysed during the same CHN run.
The magnitude of the masses on these additional components in comparison with the magnitude of the masses of aDOC and uPOC filters are presented in Figure 4. The median of the carbon mass from all the tin capsules used in the 16 CHN runs was 2(±1) µg, whilst the medians of the mass corresponding to non-acidified and acidified filter blanks were 3(±1) µg and 4(±1) µg, respectively, indicating that our method minimised contamination during acidification. On average, the acidification and handling of the filters resulted in contamination of 1(±1) µg. Nevertheless, when comparing corresponding sets of acidified and non-acidified filter blanks, the carbon masses from the acidified filters could be up to twice as large those from the non-acidified filters. Finally, the cumulative effect of all the biases and potential contamination that composed our aDOC blanks were 12(±4) µg in the productive zone and 9(±3) µg in the mesopelagic zone.
The range in carbon masses of our aDOC (Figure 5) is comparable with those from the Atlantic ocean reported by Cetinić et al. (2012): their average mass of DOC adsorption was 10.9 µg with 95% of the masses of their aDOC blanks within the range of 8.5 to 40.5 µg. Also, our aDOC values are within the findings of Abdel-Moati (1990), who reported varying amounts of DOC adsorption between 9.2 and 15.0 µg and 3.5 and 6.5 µg for eutrophic and oligotrophic waters, respectively.
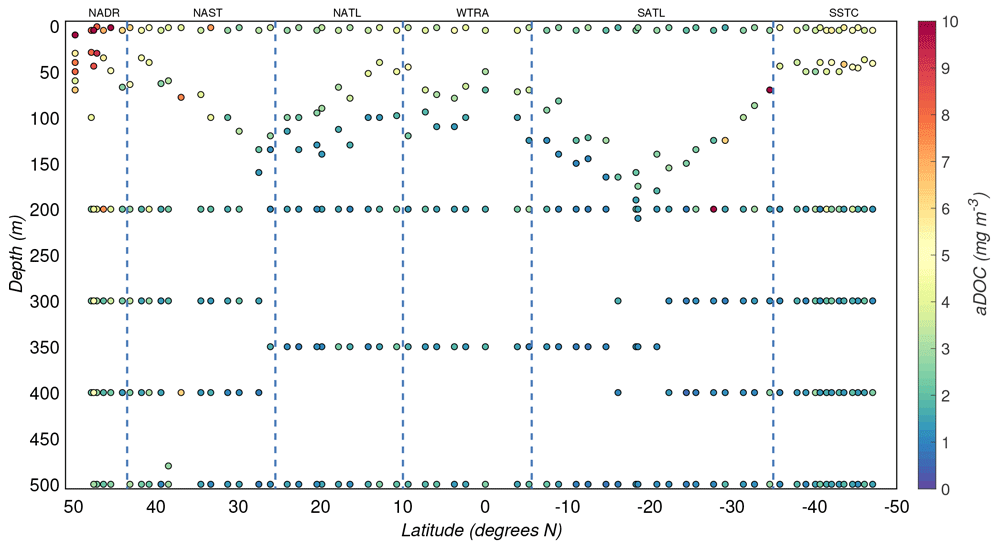
Figure 5. Depth-resolved distribution of adsorbed dissolved organic carbon (aDOC) estimates (mass of carbon on aDOC filters normalized to volume and corrected for biases) along the AMT-24 cruise.
Borders of the sampled biogeographical provinces are marked by blue vertical lines.
The aDOC concentrations (carbon mass on aDOC filters normalized by volume and corrected for biases) in the productive zone had a median value of 2(±1) mg/m3, and 1.0(±0.2) mg/m3 in the mesopelagic zone. The distribution of these concentrations was correlated with POC concentrations (r = 0.71, p<.001, Figure 9). This correlation was particularly strong in the productive zone (r = 0.87, p<.001 vs. r = 0.38, p<.001 in the mesopelagic) and in regions characterized by high surface concentrations of POC: NADR (r = 0.80, p<.001) and SSTC (r = 0.90, p<.001).
Since the filtered volumes were adjusted according to the expected POC concentration, and all GF/F filters were from a single manufacturer and treated identically during the cruise, the aDOC concentration should be relatively constant across the samples and not correlated with POC concentration. However, POC could be present on the aDOC filters, increasing the aDOC concentration due to fragmentation of particles through uPOC filters (Bishop & Edmond, 1976), perhaps due to the increased pressure differential caused by the two stacked filters (IOCCG POC Protocols). This fragmentation of larger particles from the uPOC filters into smaller particles has been observed (Wangersky, 1974) to contaminate aDOC filters and, perhaps, caused the higher aDOC values found in our study. For instance, Banoub and Williams (1972) filtered multiple samples of seawater through four stacked GF/C filters and found particles as evidence of contamination on the second filter from top to bottom. Abdel-Moati (1990) carried out a similar experiment with similar conclusions, suggesting the sum of masses from the first two filters should be used as a POC mass, while using the third filter, from top to bottom, as a true blank for aDOC. Cetinić et al. (2012) also pointed to the high variability of their aDOC values and potential contamination from the overlying uPOC filters. Finally, we note that when cells break, the DOC they contain is released in the environment and not retained on filters, potentially introducing biases in the POC determination.
If a higher than usual aDOC value is due to the contamination of an aDOC filter with particles filtered through the uPOC filter, particle loss from the uPOC filters should be higher in productive areas, thus explaining the observed strong positive correlation between POC and aDOC concentrations (Figura 9; see also Zhou et al., 2016). Assuming that particles contaminated the aDOC filters, we expect that the typical mass of organic carbon adsorbed onto our GF/F filters should be better represented by the aDOC measurements at depth, where particles are less abundant. Thus, we estimated the loss of particles from uPOC to aDOC filters by subtracting from the carbon mass of all aDOC blanks (corrected for biases), the median carbon mass of aDOC filters collected in the mesopelagic zone (≥200 m). Then, we added this difference to our POC masses and found that POC concentrations increased by 3(±4)% in the productive zone and by 0(±5)% in the mesopelagic zone.
We cannot prove which mechanism determined the correlation between aDOC and uPOC concentrations. Nonetheless, the carbon mass ratio (/), corrected for biases, ranged from 3 to 59% with medians of 9(±2)% and 12(±3)% in the productive zone and the mesopelagic zone, respectively, indicating that DOC adsorption was important.
Additional hypotheses for the observed positive correlation between aDOC and POC concentrations are that DOC and POC have a similar decreasing pattern as a function of depth in the open ocean (Dai et al., 2009) and/or that particles smaller than the nominal pore size of the filters could have passed through the uPOC filters and accidentally been retained by the aDOC filters.
3.4 Relative and total experimental uncertainties
The relative experimental uncertainty of POC concentrations, σr, (Equation 9), was on average ~12% and ~35% in the productive and mesopelagic zones, respectively. Higher σr were estimated for the mesopelagic zone where POC concentrations were lower and biases might have had a greater effect on the estimates. The resulting total experimental uncertainties of the estimates of POC concentration (σC) are shown in Figure 6. The median of σC for the productive and mesopelagic zones were 2(±2) mg/m3 and 3(±1) mg/m3, respectively.
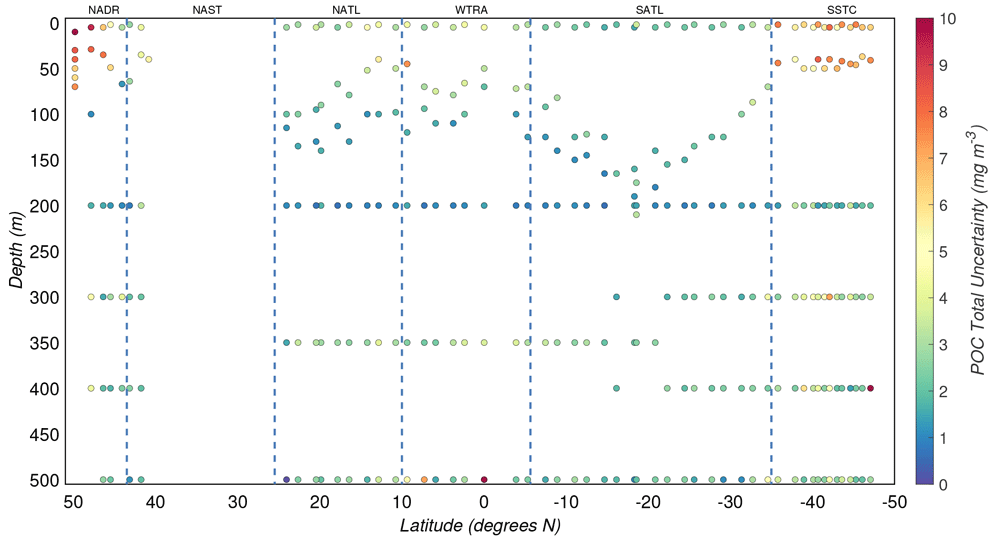
Figure 6. Depth-resolved distribution of the total experimental uncertainty associated with the estimates of particulate organic carbon (POC) concentration derived during the AMT-24 cruise.
Borders of the sampled biogeographical provinces are marked by blue vertical lines.
3.5 Uncertainty budget
Table 3 lists the sources of uncertainty that we were able to quantify and that were used to generate the uncertainty budget. The modelled uncertainty for POC is presented in Table 4 for each component, and compared to the estimated total experimental uncertainty of POC derived from the duplicate measurements. The uncertainty in volumetric measurements σC(V) typically contributed about 1% to the total uncertainty of POC concentrations in the mesopelagic zone. However, since the volume of water filtered in the productive zone was smaller, the contribution of volumetric uncertainties was greater (~2%). Overall, the contribution of this source of uncertainty was insignificant, except when POC concentration was particularly high and, as a consequence, the volume of the water sample was ~2 L.
Table 3. Sources of uncertainty contributing to the modelled uncertainty of POC (see Section 2.4.2) and total experimental uncertainty.
The median (robust standard deviation) of each uncertainty source is given for the productive (PZ) and mesopelagic (MZ) zones.
| Source | Symbol | Method of calculations | Values | Units |
|---|
| PZ | MZ |
|---|
| Volume | σV | Volume uncertainty of each sample is equal
to half graduation mark of the measuring
cylinder used to estimate the volume of each
bottle, multiplied by the number of bottles
used during a given filtration | 0.010(0.003) | 0.021(0.002) | L |
| Calibration | σMuPOC | Prediction intervals for MuPOC | 2(1) | 2(1) | μg |
| Calibration | σMaDOC | Prediction intervals for MaDOC | 2(1) | 2(1) | μg |
| Contamination | ση | Standard error of the mean of the three
acidified filters for each desiccator | 0.3(0.2) | 0.2(0.2) | μg |
| Total experimental | S | Experimental uncertainty calculated from
POC duplicates | 2(2) | 3(1) | mg/m3 |
Table 4. Uncertainty budget presenting the contributions of each source of uncertainty x that we could quantify, σC(x), relative to the total experimental uncertainty of particulate organic carbon, σC.
Median values (robust standard deviations) are given for the productive (PZ) and mesopelagic (MZ) zones.
| Source | Symbol | σC(x)/σC |
|---|
| PZ | MZ |
|---|
| Volume | σC(V) | 0.03(0.01) | 0.01(0.01) |
| Calibration | σC(M) | 0.2(0.1) | 0.10(0.08) |
| Contamination | σC(η) | 0.03(0.03) | 0.02(0.02) |
| Unquantified | σC−σC(V)−σC(M)−σC(η) | 0.8(0.1) | 0.9(0.1) |
The uncertainty due to the calibration equation σC (M), after excluding the unstable CHN runs, explained a median of 15(±11)% of the total experimental uncertainty of POC concentrations (Figure 7). Since unstable CHN runs were characterised by greater residual errors in the regression analysis, their contribution to the total experimental uncertainty was significantly higher with a median of 56(±41)%.
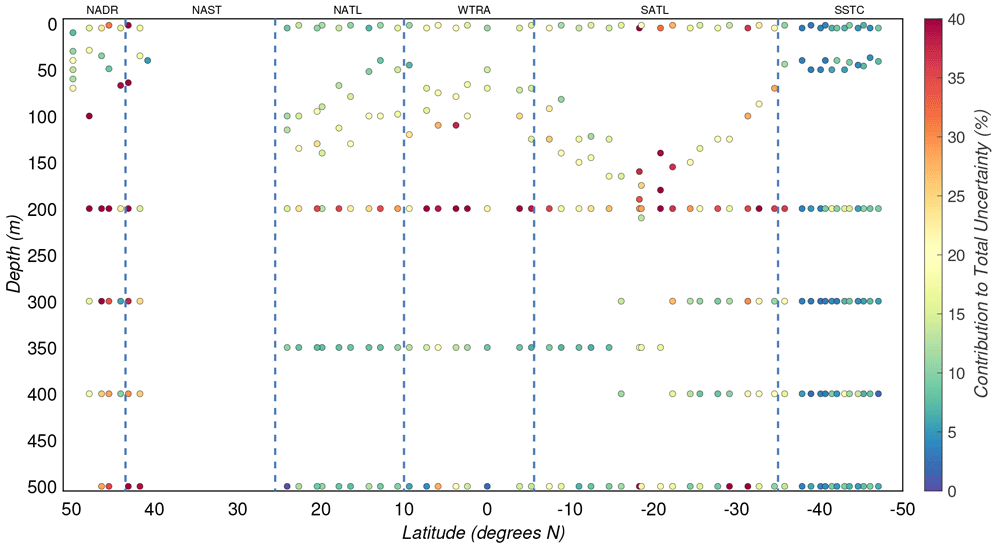
Figure 7. Percentage of the total modelled uncertainty of particulate organic carbon concentration explained by the uncertainty in the calibration.
Borders of the sampled biogeographical provinces are marked by blue vertical lines.
Our acidification method and handling during the CHN analyses allowed us to minimise the effect of contamination of POC estimates. The median contribution of this source of uncertainty, σC(η), in productive waters was 4(±4)%, while in the mesopelagic zone attained 4(±5)%.
Overall, the three sources of uncertainty described above explained only 20(±13)% of the total experimental uncertainty in POC, where medians in the productive and mesopelagic zones were 23(±13)% and 12(±9)%, respectively. Thus, other sources of uncertainty must be responsible for the relatively large and unexplained part of the estimated experimental uncertainty (Figure 8).
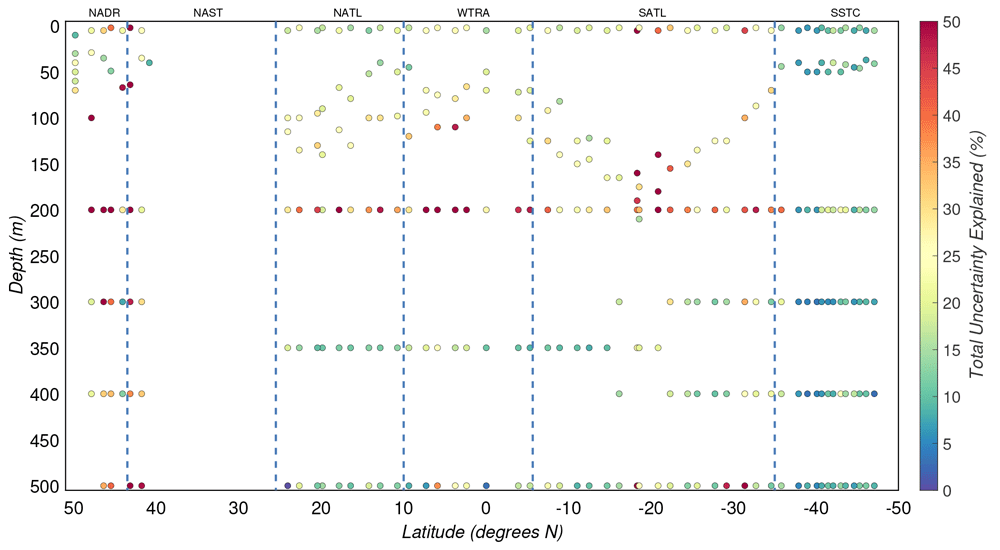
Figure 8. Percentage of the POC experimental uncertainty explained by the modelled uncertainty.
Borders of the sampled biogeographical provinces are marked by blue vertical lines.
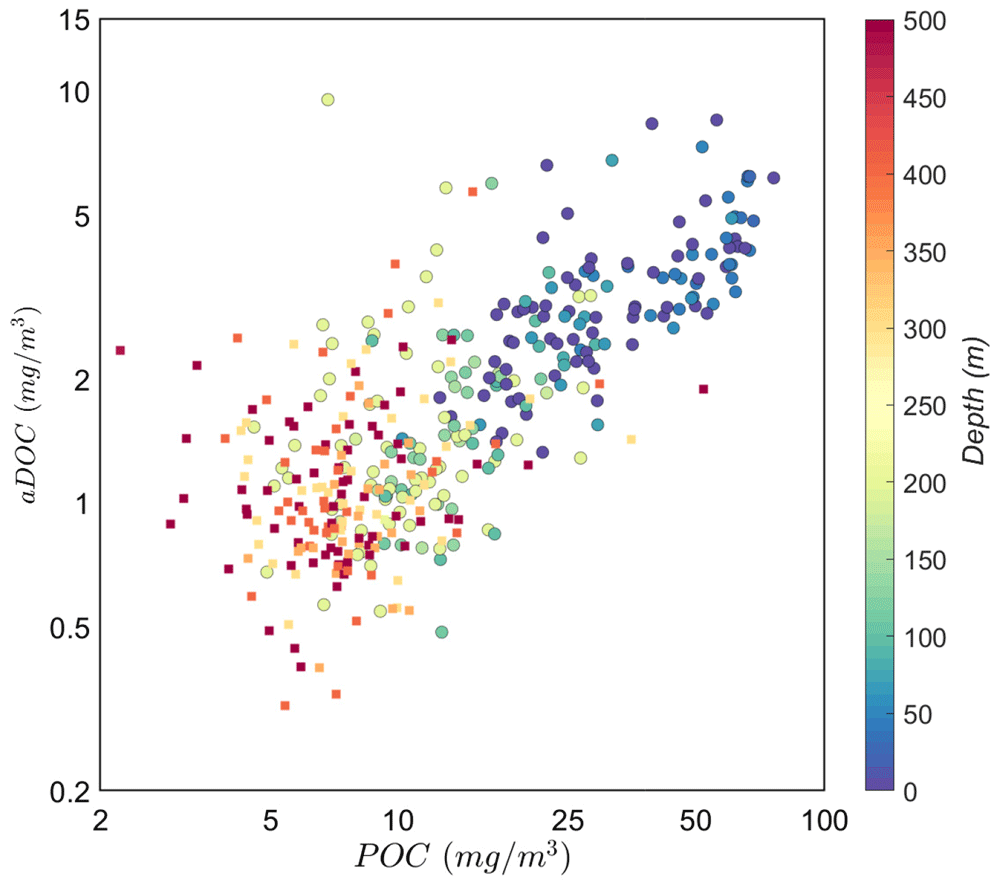
Figure 9. Correlation between aDOC and POC values.
Colours refer to sample depths. Squares represent mesopelagic samples (MZ, r = 0.38, p = 0.001). Circles represent samples in the productive zone (PZ, r = 0.87, p < 1e − 5).
3.6 Missing sources of uncertainty
In this section we discuss potential sources of uncertainty that could explain the missing part of the uncertainty budget.
3.6.1 Rare particles. Patchiness is ubiquitous in the ocean, ranging from microscale thin layers in the water column, to sub-mesoscale and mesoscale fronts, and could have contributed to our total experimental uncertainties in POC. For instance, Bochdansky et al. (2016) analysed the dynamics and abundances of particles at depth using a custom-made digital inline holographic microscope. They found that the concentration of patchy marine snow (large particles >500 µm) was 100 times higher than expected in comparison with the concentration of smaller particles. Additionally, Ohman et al. (2012) analysed the concentration and vertical distribution of suspended particulate matter and mesozooplankton at a deep-water front in the California Current System using a high resolution digital camera system. They observed that the front had a different composition of particulate matter, and it was a zone of higher marine snow particles where the volume of all size fractions of suspended particulate matter, especially organic aggregates, increased several times in comparison with the surrounding seawater. Therefore, despite we mixed samples in carboys before dispensing them to the filtration bottles, some of our duplicate filters might have captured different types of particles, i.e. rare large aggregates or (invisible) zooplankton might have appeared only on one of the two duplicate filters.
Furthermore, Wangersky (1974); Wangersky, (1976) found that a water mass in the open ocean had a homogeneous background of POC concentration, upon which occasional small patches with up to five times POC concentration were superimposed. He reported uncertainty of a single POC estimate derived from replicate 5-litre samples to be equal to 3(±1) mg/m3. Our median total uncertainty for the mesopelagic region of 3(±1) mg/m3 is comparable to these findings, suggesting that samples might have been drawn from water masses of similarly spatially variable POC concentrations.
Further experiments are required to better understand the natural heterogeneity and small-scale patchiness of seawater. Collecting marine snow particles using bottle samplers is highly challenging and time-consuming (Bochdansky et al., 2016). The oceanographic community would threfore benefit from analysing the natural heterogeneity of suspended particles and the effect of patchiness on estimates of POC concentration using optical observations from instruments such as the Underwater Vision Profiler or holographic cameras. These instruments can indeed quantify details of the in-situ vertical distribution of particles across a range of sizes (Bochdansky et al., 2016; Ohman et al., 2012). In future studies, when possible, these and other optical instruments should be used to quantify the patchiness of marine particles and its potential impacts on the uncertainty of the estimated POC concentrations. Such quantification could also inform the best strategies to collect discrete water samples in the presence of particle patchiness.
3.6.2 Contamination during filtration. The filtration system employed during this study was an open-funnel filtration set. This set up might have increased the risk of contamination by exposing the samples to particles rich in carbon (e.g., dust, ashes, ship’s engine exhaust), during sampling from the Niskin bottles and during filtration in the laboratory (Gardner et al., 2003);(IOCCG POC Sampling and Measurement Protocols).
We consider that mainly uPOC filters would have been contaminated in such a way because they were exposed to the laboratory atmosphere for longer than the underlying aDOC filters. Assuming that the uPOC filters were contaminated during filtration, their duplicate filters might have received a similar amount of contamination. Thus, atmospheric contamination of uPOC filters would result in greater differences between uPOC duplicates compared to the differences between less contaminated aDOC duplicates. Indeed, the median of the absolute differences between duplicate uPOC concentrations was 2(±2) mg/m3, which is higher than the median of the absolute difference between duplicate aDOC concentrations of 0.2(±0.4) mg/m3. This result, however, is also consistent with our previous finding that the differences between duplicate concentrations depend on POC concentrations (see Figure 2). As a consequence, we do not have enough grounds to state that differences between duplicate uPOC concentrations are higher than the differences between duplicate aDOC concentrations because of contamination of uPOC filters.
The longer a filter is exposed to the laboratory atmosphere, the more contamination it should receive. The duration of filter exposure to the laboratory atmosphere depends on the volume of seawater filtered through the filter. To further investigate contamination of uPOC filters, we analysed how differences between duplicate uPOC concentrations depended on differences between volumes of seawater filtered through two duplicate filters and found no significant correlation.
Thus, there was no evidence to suggest that the time of exposure of filters to the atmosphere was the cause of disagreement between duplicate estimates. However, analysing duplicates to determine this source of contamination might not be sufficient as duplicate estimates could have been affected by other biases or contaminants that mask a single source of contamination. Consequently, with the available data, we cannot quantify this source of contamination and its uncertainty.
For future experiments, especially when using an open-funnel setup, filtering Milli-Q water through additional blank GF/F filters may be used to quantify contamination from the laboratory atmosphere. Even better, to minimise contamination from airborne particles, it would be advisable to filter samples under a laminar flow hood or by employing a closed filtration system (Cetinić et al., 2012).
3.6.3 Storage of samples. Freezing and storing of sample filters might have also introduced some contamination. Published values suggest that the average mass of unused GF/F filters from a cruise may range from 3(±10) µg to 10(±5) µg (Cetinić et al., 2012; Menzel, 1966; Stramski et al., 2008; Wangersky, 1974). The difference between our non-acidified (3±1 µg) filter blanks and these published values might indicate that the contamination during filter storage could amount to between 0 and 7 µg. However, assuming homogeneous contamination of uPOC and aDOC filters, we would expect that this contamination would be accounted for when aDOC is subtracted from uPOC (Equation 4). To quantify uncertainties due to filter storage, we recommend preserving multiple unused GF/F filter blanks along with the samples for post-cruise analysis.
3.6.4 Collection of samples. Uncertainties can also be introduced by different operators. In our case, samples were collected by two operators, whereby one operator systematically collected samples from pre-dawn casts, while the other from noon casts. Thus, we thought that analysing duplicates pairs collected during pre-dawn and noon casts separately might give us an insight into this source of uncertainty. Due to constraints in the water budget, pre-dawn duplicates were collected from quasirandom depths, while 25 out of 28 duplicate pairs collected during noon time represented deep waters (≥400 m). For pre-dawn duplicates, the median POC concentration and the median absolute differences of duplicate POC concentrations were 9(±9) and 1(±2)mg/m3, respectively, while for noon duplicates were 9(±5) and 2(±2) mg/m3, respectively, suggesting that there was no statistical difference between these medians in the two groups of duplicates. If we take into consideration that the majority of noon duplicates were collected from deep waters (≥400 m), pre-dawn duplicates seem to be slightly more precise than noon duplicates with medians of the relative differences of duplicates collected from deep waters of 19(±10)% and 19(±31)% for pre-dawn and noon duplicates, respectively. However, there is no evidence that samples collected by different operators are biased by the operators themselves rather than by varying composition of particles at different depths. Even though the higher uncertainties that we found in the mesopelagic zone might be partially explained by the varying precision of duplicates collected by the two operators from pre-dawn and noon casts, we cannot quantify the bias and the uncertainty introduced by each operator.
3.6.5 Uncertainty model. The uncertainty model we employed in this study (Equation 10) was based on the empirical relationship we observed between duplicate differences and POC concentrations (Figure 2). Admittedly, this model is likely an approximation of the experimental uncertainty of our POC measurements. To improve the model (e.g., by adding a constant offset to it), we would need to better understand the role that each source of uncertainty plays in the total uncertainty of POC concentrations. These additional steps would ultimately allow us to then understand the extent to which each source of uncertainty is either a multiplicative or an additive term to the total uncertainty as POC, as the POC concentration in the ocean varies. Further dedicated experiments and analyses would be needed to achieve this deeper understanding.
3.7 aDOC blanks
Even though the double-filter technique employed in this study significantly increases filtration times, this procedure allowed us to collect a sample blank (i.e., aDOC filter) for each uPOC filter. Therefore, it is interesting to investigate how the uncertainty in the final POC concentrations would vary if fewer or no aDOC blanks were collected.
Some researchers avoid collecting aDOC blanks, under the assumption that by maximising the filtered volumes of seawater, uncertainties related to aDOC could be minimised (e.g., Stramski et al., 2008). Here, we can test this assumption under different scenarios by exploiting the multiple types of blanks (i.e., aDOC, non-acidified, and acidified) we have collected.
First, uPOC concentrations were higher than POC concentrations by about 13(±7)% in the productive zone and 19(±11)% in the mesopelagic zone. Biases of this magnitude have been described before (Gardner et al., 2003, and references therein). Second, by subtracting the median carbon mass of non-acidified filters (i.e., clean GF/F filter blank) from the uPOC carbon mass, we obtained uPOC concentrations (corrected for the clean GF/F filter blank) that were greater than our POC concentrations by 9(±6)% in the productive zone and 12(±9)% in the mesopelagic zone. Finally, by subtracting the median carbon mass of the acidified filters from the uPOC carbon mass, the uPOC concentrations (corrected for the acidified GF/F filter blank) were larger than our original POC concentrations by 8(±6)% in the productive zone and 10(±8)% in the mesopelagic zone. Thus, by not correcting for aDOC blanks, we would have introduced positive biases in POC concentrations of the order of 10-20%, even if we filtered up to 8 litres of seawater. It is important to realise that these results depend on the relative amount of adsorbed DOC and POC present on the filters and therefore on the volumes of water filtered and the POC concentration. One must be careful when extrapolating our conclusions to POC values determined from different sample volumes and different ocean regions.
To reduce filtration time, one could collect fewer aDOC blanks (e.g., only one deep sample per station). To quantify the potential uncertainty introduced by this method, we corrected our uPOC estimates using a single value of aDOC blank, which was determined from the median of the aDOC blanks from deep (≥200 m) stations. The resulting POC concentrations were 3(±4)% and 0(±6)% higher than the original concentrations in the productive and the mesopelagic zones, respectively. Thus, by using fewer aDOC blanks, one could significantly reduce the bias generated when not using an aDOC blank. Overall, the above exercises can guide quantitatively how many (if at all) aDOC blanks to collect, based on the level of uncertainty that one is willing to accept.
Finally, an alternative method to decrease filtration times could be to collect aDOC samples by filtering smaller amounts of water, by collecting aDOC samples separately from the uPOC ones. For example, based on an analysis of various coastal and open-ocean samples Novak et al. (2018) suggested that GF/F filters are saturated with DOC after about 0.6 liters of sample water have been filtered. Even by adopting a conservative approach and doubling this suggested DOC saturation volume one could significantly decrease filtration times at sea, while ensuring that POC values are corrected for the adsorbed DOC.
3.8 The need for a POC reference material
The accuracy of oceanographic chemical analyses is typically assessed by measuring consensus or certified reference materials (CRM). Unfortunately, at present no such CRM has been selected by the oceanographic community for POC analyses (IOCCG POC Sampling and Measurement Protocols). CRMs exist that potentially might be used to represent organic matter in the ocean, e.g., NIST Buffalo River Sediment RM 8704 (National Institute of Standards and Technology), but their precise bio-organic elemental composition has not been determined, which prevents one from assessing how representative they are of open-ocean pelagic particulate matter (National Research Council, 2002). In addition, these CRMs are mainly comprised of marine sediments, rich in aluminosilicates and quartz, but with no pelagic opal and carbonate matrices, thus misrepresenting the complex matrix associated with open-ocean pelagic POC samples and potentially introducing artefacts in the accuracy assessment (National Research Council, 2002). The Committee on Reference Materials for Ocean Science of the US National Research Council recommended that a consensus or certified reference material for POC representative of open-ocean particulate matter could be obtained by mixing cultures from a diatom, a dinoflagellate, and a coccolithophore (National Research Council, 2002). Unfortunately, as of today, no CRM for POC analyses has been developed and, as a consequence, POC analyses are difficult to compare over time, among groups, and when different analytical protocols are used. Future work to improve POC determinations should focus on agreeing upon and producing a certified or consensus reference material. Finally, intercomparison exercises are also needed to minimise uncertainties arising from all sample collection and processing steps before the CHN analysis. Dedicated funding and an international effort are needed to achieve these two crucial objectives.
4 Conclusions
In this study we the quantified experimental uncertainties of our POC concentrations and compared them with modelled uncertainties based on assumed sources of uncertainties. We found that the total experimental uncertainty of our POC estimates varied with depth and with POC concentration and was ~12% and ~35% in the productive and in the mesopelagic zones, respectively. However, we could not identify all the different sources of uncertainty associated with POC concentrations, and our modelled uncertainty could explain only ~19% of the total experimental POC uncertainty. Further work is required to identify the unexplained portion of the modelled uncertainty.
Nevertheless, this study improved our understanding of the limitations of the method and of some of the stages of sample collection and processing that contributed to the variability of our results. Further experiments would be required to fully understand the uncertainty budget of POC estimates. This understanding would allow us to concentrate our efforts on those parts of the methodology that are more prone to introduce uncertainties and to develop a better-informed protocol to improve comparability of POC estimates across different studies.